Cancer cells can shrug off powerful drugs without ever changing their DNA, and a new review article argues that the explanation may lie in a level of biology that drug design has largely ignored: the constantly shifting three-dimensional shapes of proteins. Writing in the British Journal of Cancer, biophysicists Ruth Nussinov and Hyunbum Jang propose that dynamic conformational heterogeneity—the fact that proteins exist as fluctuating populations of many shapes rather than a single static structure—acts as a fundamental driver of adaptive phenotypic plasticity and therapeutic resistance. The authors present this as a conceptual framework rather than an established mechanism, and they state plainly that the role they propose for transient conformational substates in plasticity remains a hypothesis awaiting direct experimental validation.
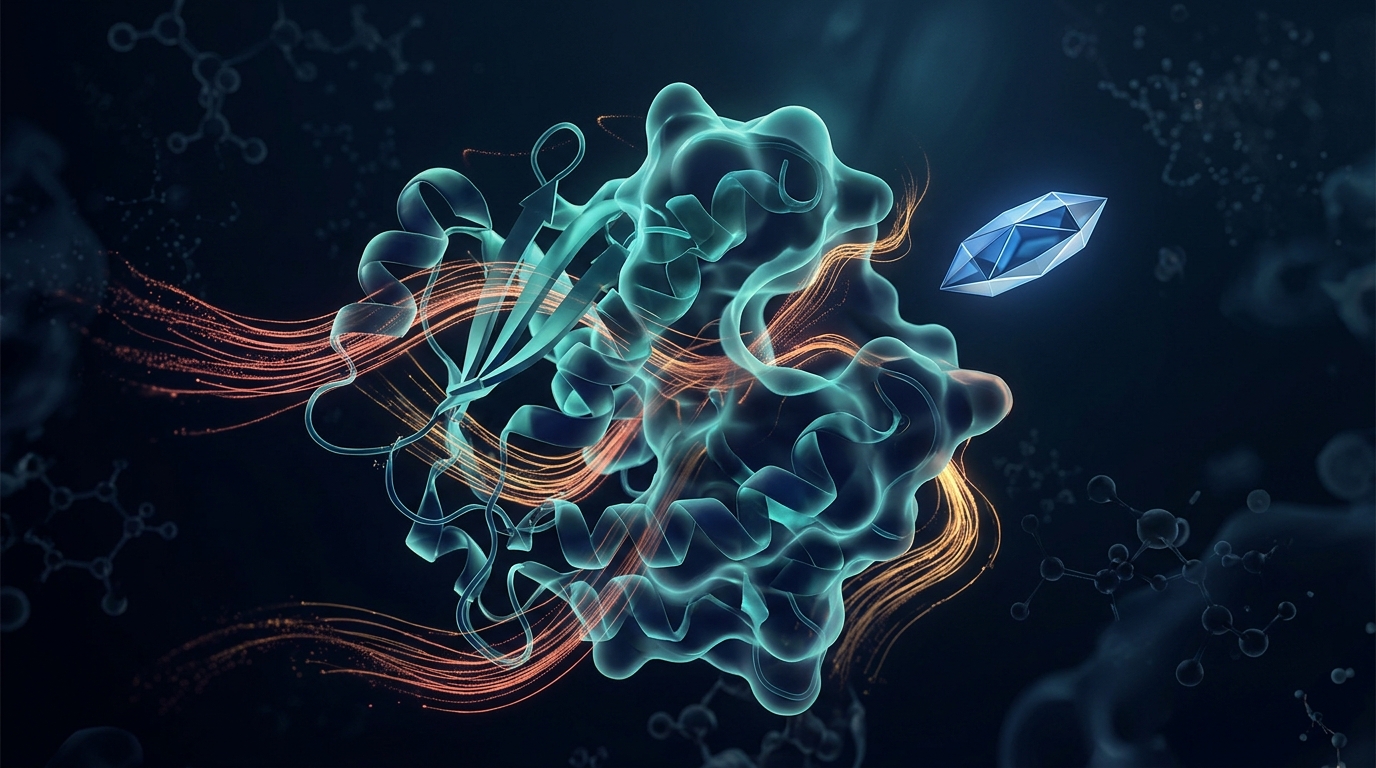
The core idea challenges a long-standing habit in pharmacology. Structure-based drug design typically relies on a single, static crystal structure of a target protein, yet the authors argue that such a structure is constrained by the crystal lattice and influenced by solutes. At physiological temperatures inside a cell, the conformation captured in the crystal may be only sparsely populated. Effective drugs, in their account, often work through conformational selection: they bind one specific shape within the ensemble and shift the equilibrium toward that state, followed by minor backbone and side-chain induced-fit optimization. Resistance arises when mutations or altered cellular conditions redistribute the ensemble so that the protein spends more time in shapes the drug cannot stably recognize.
Proteins, in this view, are not binary switches. They bounce around a rugged free-energy landscape dotted with hills and valleys, jostled by thermal energy that makes atoms collide billions of times per second. Deep valleys correspond to major conformational states with long residence times, separated by high kinetic barriers on the microsecond-to-millisecond scale. Within those basins sit metastable substates separated by much lower barriers, transitioning on picosecond-to-nanosecond timescales. The authors define a kinetic bridging substate as a transient, metastable conformational substate that connects neighboring free-energy minima along a productive transition pathway. Unlike a transition state, which sits at the top of an energy hill with essentially zero occupancy, a bridging substate is a populated local minimum with a finite lifetime and is, in principle, experimentally observable through advanced NMR spectroscopy, single-molecule kinetics, and molecular dynamics simulations.
From these physical premises the authors build a hypothesis about how cells inherit drug-tolerant states without new mutations. Rather than relying on static molecular marks, they propose that cell state memory is driven by the dynamic distribution of protein conformational ensembles that epigenetic marks modulate. A cancer cell, they suggest, can hop into a pre-existing but low-frequency conformational substate that happens to be drug-tolerant, bridging statistical mechanics and clinical oncology. Because returning to the original state requires crossing the same kinetic barriers, previous occupancy can bias subsequent transitions, producing an apparent conformational memory. The authors are careful to distinguish scales here: conformational memory biases molecular transitions over short relaxation timescales, while chromatin modifications stabilize transcriptional programs over longer durations, including across cell divisions.
The framework also assigns a specific role to classical epigenetics and genetics. Aberrant epigenetic changes such as altered DNA methylation and histone acetylation often occur at earlier, pre-malignant stages, changing expression levels—overexpressing oncogenes and underexpressing repressors—and creating a permissive environment that promotes stem-like plasticity. Epigenetically driven states, the authors argue, often precede and set the stage for the evolution of specific mutational subclones. Later, feedback loops emerge in which genetic mutations drive further epigenetic alterations, as in IDH-driven hypermethylation, and early driver mutations in genes such as DNMT3A, TET2, EZH2, and IDH1/2 can rewrite the cell’s epigenetic landscape. In their formulation, epigenetics and genetics shape the landscape, conformational heterogeneity provides the molecular engine, and phenotypic plasticity is the adaptive process that generates intratumor heterogeneity and resistance.
Overexpression enters the model as a dosage-driven shift in the functional ensemble. Changes in the total number of protein molecules available alter the absolute population of the active conformation even if the relative propensity stays the same. In a linear system, a protein that spends only one percent of its time in an active ON state would double its ON signal if its concentration doubled. But biological systems such as Ras/ERK and PI3K/AKT/mTOR are often sigmoidal and ultrasensitive, so doubling the concentration could flip a threshold-dependent switch from negligible activity to saturation, making downstream targets constitutive. Massive dosage through high-level amplifications of genes such as HER2 and MYC can saturate the proteins responsible for negative feedback, including dual-specificity phosphatases and Sprouty proteins that normally downregulate Ras/ERK signaling. Genetic mutations act differently, lowering energy barriers or changing the equilibrium constant between states, but both routes converge on an ensemble population shift—an allosteric mechanism—that locks downstream signaling into a constitutive ON state.
This lens reshapes how the authors interpret metastasis. Metastatic cells do not show an increased genetic mutational load compared with the primary tumor, and primary tumors require the accumulation of multiple mutations, which is why their emergence can take years. Metastasis, by contrast, can be fast—possibly weeks or months—and the authors argue it is increasingly viewed as epigenetics driven, relying on rapid changes in DNA methylation and histone acetylation along with gene copy number variations, specifically oncogene duplications and repressor deletions. They note exceptions: metastatic tumors can carry resistance mutations such as ESR1 and AR in hormone-treated breast and prostate cancers, EGFR T790M in non-small cell lung cancer treated with first- or second-generation EGFR tyrosine kinase inhibitors, and KIT V654A in imatinib-treated gastrointestinal cancer. Metastatic cells also often show an increase in copy number alterations, which drive large-scale gene expression changes more potently and durably than epigenetic alterations alone.
The review grounds the argument in glioblastoma, an aggressive and lethal brain cancer characterized by extensive genomic instability. For IDH-wildtype glioblastoma, the combined gain of chromosome 7 and loss of chromosome 10 is described as a pathognomonic feature. Chromosome 7 gain increases copy numbers and overexpression of oncogenes including EGFR, CDK6, MET, and BRAF, while chromosome 10 loss eliminates tumor suppressor genes such as KLF6, ANXA7, and PTEN. Homozygous deletion of the CDKN2A and CDKN2B locus on chromosome 9p21 is frequently observed, disrupting the p16INK4a and p14ARF cell-cycle regulatory proteins, and amplification of CDK4 and MDM2 within the 12q14-15 region is often driven by extrachromosomal DNA. Copy number alterations are nearly universal in aggressive cancers, occurring in up to 99 percent of certain high-grade tumors such as aggressive meningiomas, and CNA burden has been offered as a predictor of aggression.
What follows for therapy is a two-level strategy. Because conformational heterogeneity itself is not directly druggable, the authors argue that treatments must target the molecular and network-level mechanisms through which it promotes plasticity. At the molecular level, therapies should stabilize pharmacologically favorable conformational states using allosteric ligands, molecular glues, PROTACs, or epigenetic modifiers, while promoting selective degradation of pathogenic conformers. Molecular glues stabilize a sparse pre-existing population of protein-protein interfaces, stabilizing ternary complexes that induce proximity and a favorable orientation for degradation, and allosteric PROTACs are being developed for targets including TEAD of the Hippo pathway and the SH2 domain of STAT3. At the systems level, target-node selection should prioritize not only mutational variants but the regulatory architecture that sustains plasticity—pathway cross-links, negative feedback circuits, and transcriptionally controlled signaling nodes—so that overexpressing cells cannot simply redistribute signaling through alternative pathways as resistance evolves.
The authors acknowledge that transcription and epigenetic factors are difficult targets: they often lack defined, stable small-molecule binding pockets, act through broad protein-protein or protein-DNA interfaces, are frequently intrinsically disordered, and reside in the nucleus, so drugs must cross the nuclear membrane. They also caution that the relative contribution of conformational memory versus rigid genetic tags varies across cancer types depending on the baseline epigenetic stability of the tissue of origin, and that crowded biomolecular condensates, while stabilizing drug binding, challenge drug delivery through increased viscosity and altered diffusion kinetics. The proposed role of kinetic bridging substates in adaptive plasticity, they stress, awaits direct experimental validation through methods such as FRET, rapid-mixing NMR, or perturbation assays. If validated, the framework would push oncology further from single-drug, single-structure thinking toward adaptive, system-level drug combinations aimed at the dynamic populations of shapes that cancer cells exploit to survive.
Subject of Research: A review proposing protein conformational heterogeneity as a proposed mechanism of adaptive phenotypic plasticity and therapeutic resistance in cancer.
Article Title: Conformational heterogeneity: a fundamental mechanism of adaptive phenotypic plasticity and therapeutic resistance
Article References: Nussinov, R., & Jang, H. (2026). Conformational heterogeneity: a fundamental mechanism of adaptive phenotypic plasticity and therapeutic resistance. British Journal of Cancer. https://doi.org/10.1038/s41416-026-03649-x
Image Credits: AI Generated
DOI: 10.1038/s41416-026-03649-x
Keywords: conformational heterogeneity, protein ensembles, drug resistance, phenotypic plasticity, epigenetics, copy number alterations, glioblastoma, conformational selection, allosteric drugs, metastasis, kinetic bridging substates, cancer therapy
News Source: Nathaniel Bowman. (November 3, 2026). Cancer’s shape-shifting proteins may explain why drugs stop working. Scienmag.