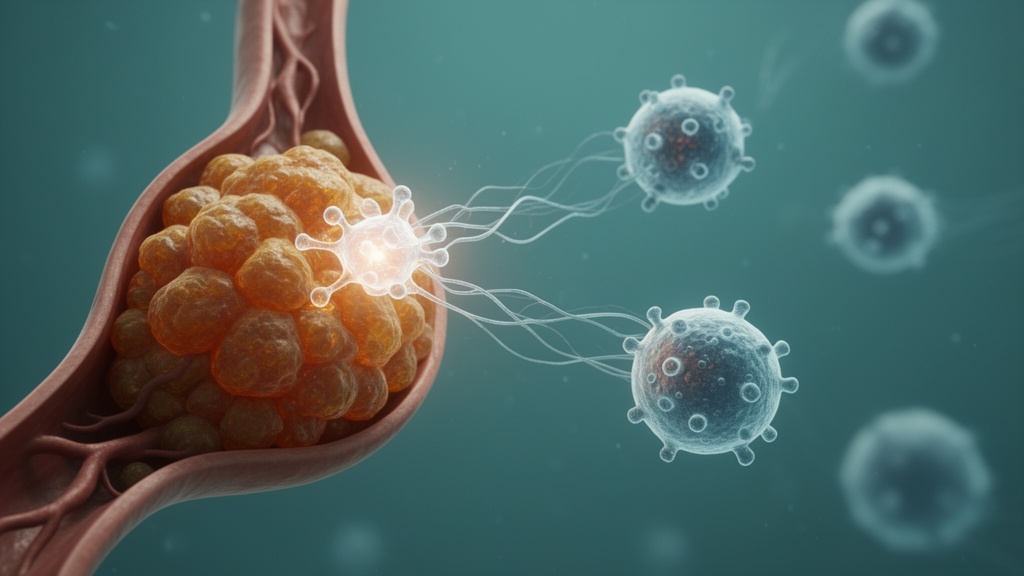
Intrahepatic cholangiocarcinoma, an aggressive cancer arising from the bile ducts inside the liver, is notorious for its poor response to immunotherapy. A new study published in Journal of Translational Medicine points to one reason why: a tumor-produced protein called stratifin, or SFN, appears to help the cancer silence natural killer cells, the immune cells that can destroy tumor cells without prior antigen presentation. The researchers report that tumors with high SFN expression showed reduced intratumoral NK-cell abundance and impaired NK-cell functional signatures, and that SFN expression was associated with poor patient outcomes and a negative relationship with immune infiltration. Together, the findings outline a tumor-intrinsic mechanism that reshapes communication between cancer cells and NK cells in this hard-to-treat malignancy.
To identify the regulator, the team integrated bulk RNA sequencing, single-cell RNA sequencing, proteomics, tissue microarrays, and external patient cohorts to characterize immune remodeling in ICC and prioritize candidate drivers. Stratifin emerged as a leading candidate because it was highly expressed in ICC tumor cells, tracked with worse prognosis, and inversely related to immune infiltration. The authors then moved from correlation to causation using ICC cell lines, NK-cell co-culture systems, and murine ICC models, assessing NK-cell infiltration and activity by flow cytometry, immunofluorescence, immunohistochemistry, and functional assays.
The functional experiments centered on the mIC-23 orthotopic ICC model, in which tumors grow in their natural liver environment. When the researchers knocked down SFN in tumor cells, NK-cell infiltration and effector activity increased. The reverse experiment sealed the logic: when NK cells were depleted, the antitumor effect of SFN knockdown was attenuated, indicating that NK cells are required for much of the benefit. In other words, SFN appears to act as a molecular brake that tumors apply to their own microenvironment, and removing the brake reawakens the innate immune arm that ICC has largely escaped.
To trace how SFN exerts this effect, the team used CellChat-based ligand–receptor inference to nominate candidate tumor–NK signaling axes, then tested them with co-culture, genetic rescue, and receptor-blockade assays. The evidence converged on the NECTIN2/TIGIT axis: NECTIN2, a ligand displayed on the tumor cells, engages TIGIT, an inhibitory receptor on NK cells, damping their cytotoxic response. Mechanistic work with co-immunoprecipitation, ubiquitination assays, ChIP-qPCR, and dual-luciferase reporter assays then revealed the circuitry upstream of NECTIN2. Stratifin was found to interact with PAK1 and was associated with reduced TRIM21-related K48-linked ubiquitination of PAK1, thereby increasing PAK1 stability. Stabilized PAK1 fed into AKT/GSK-3β signaling, which contributed to the activation of MYC, a well-known transcriptional amplifier of tumor growth. MYC, in turn, participated in the transcriptional regulation of NECTIN2, completing a chain that runs from a tumor-cell protein all the way to the inhibitory signal that disarms NK cells.
The translational payoff came from combining therapies. In the mIC-23 orthotopic model, chemotherapy combined with PD-1 and TIGIT blockade enhanced antitumor responses and NK-cell activity, suggesting that attacking the TIGIT checkpoint alongside conventional treatment can restore innate immune pressure on SFN-high tumors. Exploratory retrospective analyses in patient data also suggested a potential association between elevated SFN and reduced immunotherapy responsiveness, raising the possibility that SFN could serve as a marker for selecting patients likely to benefit from TIGIT-targeted combinations. The authors caution that these retrospective findings are exploratory and that the therapeutic results come from preclinical models, so the combination will require further clinical evaluation before it can inform practice.
The study’s significance lies in how it connects several layers of biology into a single, testable model. Tumor-intrinsic SFN signaling, acting through a PAK1–MYC-associated axis, drives NECTIN2 expression on tumor cells, which in turn engages TIGIT on NK cells and suppresses their function. This provides mechanistic insight into why NK-cell dysfunction is so pervasive in ICC and why immune checkpoint blockade alone has delivered limited benefit for many patients. It also reframes stratifin from a passive prognostic marker into an active participant in immune evasion, one whose expression could, in principle, be measured to stratify patients or targeted directly to sensitize tumors to NK-cell attack.
Subject of Research: Tumor-intrinsic stratifin regulation of NECTIN2–TIGIT-mediated NK-cell suppression in intrahepatic cholangiocarcinoma
Article Title: Tumor-intrinsic stratifin contributes to NECTIN2–TIGIT-mediated NK-cell suppression and therapeutic vulnerability in intrahepatic cholangiocarcinoma
Article References: Sun, G., Wang, Z., Gan, W., Yang, Z., Gao, X., Han, Z., Wang, C., Zhang, C., Hu, B., Zhou, C., Yi, Y., & Qiu, S. (2026). Tumor-intrinsic stratifin contributes to NECTIN2–TIGIT-mediated NK-cell suppression and therapeutic vulnerability in intrahepatic cholangiocarcinoma. Journal of Translational Medicine. https://doi.org/10.1186/s12967-026-09057-2
Image Credits: AI Generated
DOI: 10.1186/s12967-026-09057-2
Keywords: intrahepatic cholangiocarcinoma, stratifin, natural killer cells, NECTIN2, TIGIT, PAK1, MYC, immunotherapy, tumor microenvironment, immune evasion, Tumor-intrinsic, contributes
News Source: Nathaniel Bowman. (October 23, 2026). Tumor protein stratifin blinds NK cells in bile duct cancer. Scienmag.