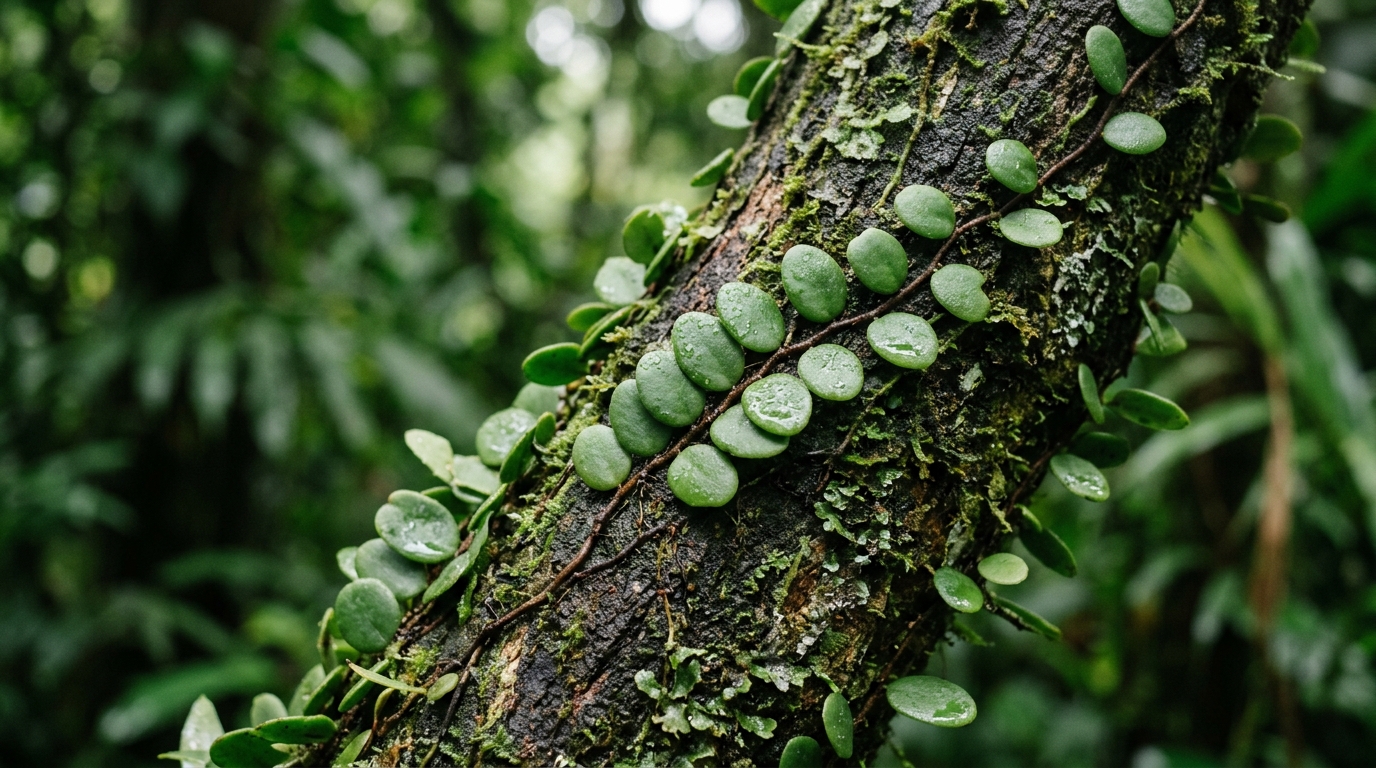
A new computational investigation has identified eight bioactive compounds within the dragon scale fern, scientifically known as Pyrrosia piloselloides, that exhibit potential for therapeutic interaction with proteins implicated in prostate cancer. Prostate cancer remains a critical health concern globally, ranking as the second most commonly diagnosed cancer among men and the fifth leading cause of cancer-related mortality. Despite clinical advancements, intrinsic or acquired resistance to common chemotherapeutic agents such as docetaxel and bicalutamide presents a persistent challenge. The researchers employed a comprehensive, integrated in silico workflow—encompassing network pharmacology, molecular docking, and molecular dynamics—to explore whether compounds from this traditional medicinal plant might offer a new framework for targeting proteins associated with the disease. This computational study, which does not constitute a clinical trial or experimental confirmation, provides a hypothesis-generating platform for prioritizing these botanical candidates for future laboratory validation.
To establish a reliable set of study subjects, the investigators began by gathering 59 unique bioactive compounds identified in previous gas chromatography-mass spectrometry (GC-MS) profiles of aqueous and methanolic leaf extracts of Pyrrosia piloselloides. These compounds were subjected to a rigorous screening process designed to evaluate drug-likeness and predicted oral bioavailability. Using established tools such as SwissADME, Molsoft, and the Traditional Chinese Medicine Systems Pharmacology (TCMSP) database, the team applied stringent criteria, including a molecular weight of less than 500 Da, defined hydrogen bond donor and acceptor counts, and specific thresholds for drug-likeness indices. After the elimination of compounds that did not meet these pharmacokinetically favorable criteria, eight candidates remained for further analysis: 6-hydroxymethaqualone, 2,6,10,14,18,22-tetracosahexaene, ergost-8(14)-en-3beta-ol, stigmasterol, chondrillast-7-enol, stigmastan-3,5,22-trien, 7-dehydrodiosgenin 3-acetate, and beta-sitosterol acetate.
The study then sought to determine if these phytochemicals could target key proteins involved in prostate cancer progression. By utilizing target prediction platforms including SwissTargetPrediction and PharmMapper, and cross-referencing these findings with disease-associated proteins identified in databases such as GeneCards, OMIM, and DisGeNET, the researchers identified 32 proteins that intersected between the plant compounds and prostate cancer. Subsequent topological analysis of the protein-protein interaction network—constructed via the STRING database and analyzed using Cytoscape software—allowed the team to isolate a primary network of 30 interconnected proteins. Through the application of centrality parameters such as degree, betweenness, closeness, and the maximum neighborhood component, the researchers identified nine hub proteins that likely represent the core targets for these compounds: ESR1, MYC, CTNNB1, PPARG, AKT1, STAT3, BCL2, AR, and ERBB2.
Functional enrichment analysis provided further insight into the biological significance of these targeted proteins. Gene Ontology (GO) assessment revealed that the targets were enriched in molecular functions related to nitric oxide synthase regulator activity and nuclear estrogen receptor binding. Biological process analysis, covering 1,000 terms, highlighted epithelial cell proliferation, hormone responses, and cell signaling regulation as the most highly enriched processes, consistent with the known progression factors of prostate cancer. Furthermore, KEGG pathway enrichment analysis identified 161 pathways, with eight demonstrating strong relevance to prostate cancer biology, including the PI3K–Akt signaling pathway, endocrine resistance, and the ErbB signaling pathway. These results suggest that Pyrrosia piloselloides compounds might engage in a multifaceted regulatory role rather than acting through a single molecular mechanism, providing a broad, multi-target approach to potential therapeutic intervention.
The team next employed molecular docking with AutoDock Vina to predict the binding affinity of the eight compounds against the nine identified core protein targets. The study protocol was validated by re-docking co-crystallized native ligands and ensuring that the root mean square deviation (RMSD) remained below 2.0 Å. The resulting docking simulations revealed that all eight plant-derived compounds interacted with the target proteins. Notably, 7-dehydrodiosgenin 3-acetate demonstrated the most negative docking scores across multiple targets, including a significant affinity for ERBB2 at −11.85 kcal/mol, AKT1 at −9.49 kcal/mol, and BCL2 at −9.38 kcal/mol. Stigmasterol also showed potent interactions, specifically for PPARG at −9.63 kcal/mol and AR at −9.31 kcal/mol. These variable affinities suggest that specific phytochemicals may have preferred interaction profiles among the array of critical prostate cancer drivers.
To investigate the dynamic behavior and persistence of these interactions, the researchers performed 100 ns molecular dynamics simulations on the most promising protein-ligand complexes using the OpenMM platform. By tracking the RMSD, RMSF, and the radius of gyration, the team assessed whether the ligand binding disrupted the global structural integrity of the target proteins. The simulations for the ERBB2–6-hydroxymethaqualone, PPARG–Stigmasterol, and BCL2–7-dehydrodiosgenin 3-acetate complexes indicated stable structural behavior. Following an initial equilibration, RMSD values remained relatively stable for the duration of the 100 ns runs. While some residue fluctuations were noted in the RMSF profiles, these were primarily localized to flexible loop or terminal regions, and the radius of gyration remained constant throughout the simulation, supporting the conclusion that the global compactness of the target proteins was preserved upon ligand binding.
Further binding free energy analysis was conducted using the Molecular Mechanics/Generalized Born Surface Area (MM/GBSA) method to quantify the stability of these interactions. The ERBB2–6-hydroxymethaqualone complex yielded the most favorable binding free energy (ΔG) at −45.57 kcal/mol. This was followed by the PPARG–Stigmasterol complex at −41.52 kcal/mol and the BCL2–7-dehydrodiosgenin 3-acetate complex at −23.35 kcal/mol. Across all three complexes, the binding affinity was largely driven by favorable van der Waals interactions, which are characteristic of the predominantly hydrophobic nature of the phytochemicals. While electrostatic contributions were marginal, the positive contribution of solvation free energy partially offset these favorable gas-phase interactions, ultimately resulting in stable negative binding free energies that underscore the potential stability of these compound-target pairings.
The study also included a pharmacokinetic evaluation using tools to forecast absorption, distribution, metabolism, excretion, and toxicity (ADMET) profiles. The findings indicated that the test compounds generally exhibited strong predicted human intestinal absorption, often exceeding that of the reference drug flutamide. Regarding metabolic pathways, most compounds were predicted to be CYP3A4 substrates, a common metabolic route for several clinically approved anticancer agents. Notably, the majority of the selected compounds did not demonstrate predicted inhibition of major cytochrome P450 isoforms, suggesting a potentially lower likelihood of problematic drug-drug interactions, although this prediction requires empirical confirmation. Regarding potential safety, the compounds were broadly classified as non-hepatotoxic, and acute oral toxicity predictions were generally favorable, though specific variations in mutagenic and toxic potential were observed among the compounds, particularly with 6-hydroxymethaqualone.
It is essential to interpret these results within the context of the study’s in silico methodology. While the integration of network pharmacology and molecular simulations offers a powerful computational framework, it cannot fully replicate the complex biological systems of the human body. The study’s reliance on existing databases and target prediction algorithms entails inherent limitations, including the possibility of false-positive or false-negative results. Furthermore, the binding free energies calculated via MM/GBSA should be viewed as relative estimates rather than absolute values, as explicit entropic contributions were not accounted for in this analysis. The identification of these candidates should therefore be treated as a foundation for hypothesis-driven research, with experimental laboratory and, eventually, clinical validation remaining necessary to confirm actual target engagement, biological efficacy, and clinical safety.
In conclusion, the study successfully applied an integrated computational approach to map potential interactions between phytochemicals in Pyrrosia piloselloides and prostate cancer-related targets. By identifying 7-dehydrodiosgenin 3-acetate, stigmasterol, and 6-hydroxymethaqualone as compounds with favorable binding profiles, the researchers have delineated specific targets—including ESR1, MYC, CTNNB1, PPARG, AKT1, STAT3, BCL2, AR, and ERBB2—that warrant further scrutiny. These pathways, particularly the PI3K-Akt and ERBB signaling networks, are heavily implicated in cancer cell proliferation and survival. While these findings do not establish a therapeutic treatment, they provide a systematic, data-backed rationale for prioritizing these specific plant-derived compounds for future experimental studies to investigate their translational potential in the management of prostate cancer.
Subject of Research: Computational identification of potential anticancer compounds from Pyrrosia piloselloides targeting prostate cancer proteins.
Article Title: Computational network pharmacology and molecular simulations to explore the potential mechanisms of Pyrrosia piloselloides compounds against prostate cancer
Article References: Havid, H., Ysrafil, Y., Fatmaria, F., Musyaffa, M. R., Sinuhaji, M. S. C., & Pratika, R. A. (2026). Computational network pharmacology and molecular simulations to explore the potential mechanisms of Pyrrosia piloselloides compounds against prostate cancer. Journal of the Egyptian National Cancer Institute, 38(1), Article 83. https://doi.org/10.1186/s43046-026-00421-3
Image Credits: AI Generated
DOI: 10.1186/s43046-026-00421-3
Keywords: Pyrrosia piloselloides, prostate cancer, network pharmacology, molecular docking, molecular dynamics, MM/GBSA, 7-dehydrodiosgenin 3-acetate, stigmasterol, 6-hydroxymethaqualone, ESR1, MYC, CTNNB1, PPARG, AKT1, STAT3, BCL2, AR, ERBB2, in silico study
News Source: Nathaniel Bowman. (November 16, 2026). Computational Study Identifies Potential Pyrrosia Piloselloides Compounds Against Prostate Cancer. Scienmag.