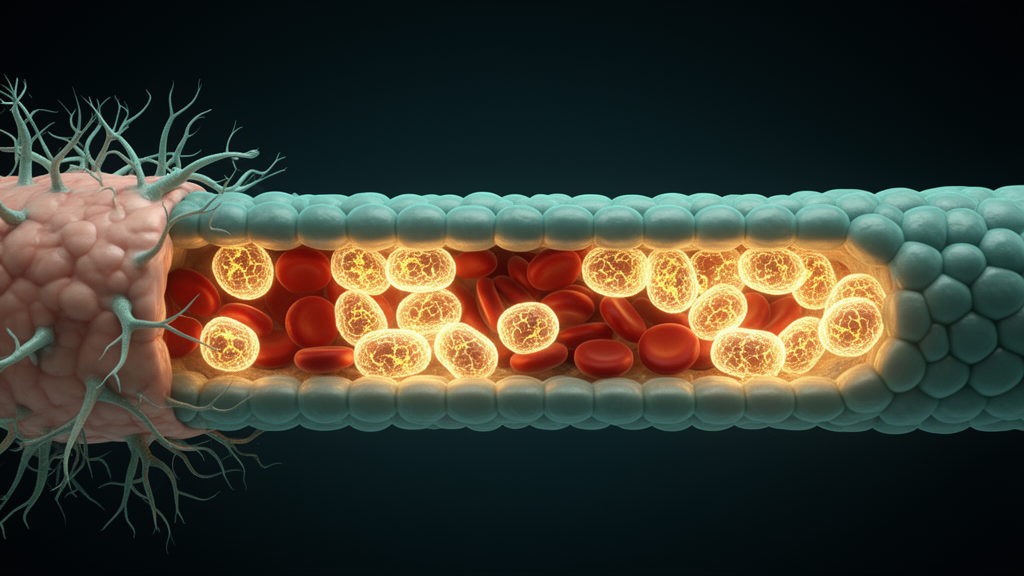
Repetitive mild traumatic brain injury (rmTBI) is increasingly recognized as a condition in which the timing of repeated impacts matters as much as the impacts themselves. A new narrative review published in Frontiers in Molecular Neuroscience argues that when concussive or subconcussive blows recur before the brain has fully recovered, mitochondria—the energy hubs of neurons and vascular endothelial cells—undergo a profound metabolic remodeling that may drive blood–brain barrier (BBB) dysfunction and set the stage for chronic neurodegeneration. The review, led by Lingjuan Gao and colleagues at Jiangyin Clinical College of Xuzhou Medical University, synthesizes evidence across the traumatic brain injury (TBI) spectrum to propose that recurrent metabolic stress, redox imbalance, inflammatory signaling, and impaired mitochondrial quality control converge on the neurovascular unit, the multicellular structure that maintains the BBB.
Unlike a single severe TBI, which causes immediate and extensive tissue necrosis, rmTBI imposes a sustained and insidious insult on mitochondria through repeated mechanical stress. The review emphasizes that mitochondrial dysfunction after rmTBI is not merely a matter of energy depletion but a complex pathological process involving structural damage, dynamic imbalance, and profound remodeling of the metabolite profile. This process, the authors argue, constitutes the core driver of the transition from acute injury to chronic neurodegeneration, including conditions relevant to chronic traumatic encephalopathy (CTE).
The biomechanical basis of rmTBI centers on shear and stretching forces generated during head acceleration-deceleration or rotational movements. Because cerebral blood vessels differ in density and stiffness from the surrounding brain parenchyma, shear forces preferentially act on tight junctions between endothelial cells—proteins such as claudin-5 and occludin—and on the basement membrane, compromising the structural basis for vasogenic edema. Stretching forces simultaneously disrupt the cytoskeleton, impair axonal transport, and activate mechanosensitive ion channels such as Piezo1, triggering early calcium influx. This calcium overload further disrupts mitochondrial oxidative phosphorylation and precipitates an energy crisis, placing neurons in an energy-vulnerable state.
At the mitochondrial level, the review details how repetitive shear and stretching forces are directly transmitted to the intracellular environment, leading to physical disruption of the mitochondrial membrane and disorganization of cristae architecture. Excessive calcium uptake via the mitochondrial calcium uniporter triggers sustained opening of the mitochondrial permeability transition pore, resulting in membrane potential collapse, uncoupling of the respiratory chain, and release of apoptotic factors such as cytochrome c. Concurrently, electron transport chain complexes are significantly suppressed: decreased activity of Complex I and Complex IV in the cerebral cortex and hippocampus following rmTBI leads to increased electron leakage and reduced ATP synthesis. The ATP/ADP/AMP ratio becomes severely imbalanced, and phosphocreatine, an immediate energy buffer, is rapidly depleted.
Beyond structural and bioenergetic dysfunction, the review highlights systemic remodeling of the mitochondrial metabolite profile. Reduced ATP synthesis increases AMP relative to ATP and activates AMPK through upstream kinases including LKB1 and CaMKK2. Acute AMPK activation can be adaptive by conserving energy and supporting mitochondrial quality control, but under persistent energy failure, dysregulated AMPK-mTOR signaling, inadequate autophagic flux, oxidative stress, and pro-apoptotic pathways may together contribute to injury. The authors caution that AMPK activation should not be interpreted as intrinsically harmful or protective; its consequence depends on duration, cellular context, and the capacity to restore bioenergetic homeostasis.
Among the key metabolites, succinate accumulation can increase mitochondrial reactive oxygen species (ROS) through reverse electron transport and can influence HIF-1alpha and inflammatory pathways. NAD+ depletion may limit sirtuin-dependent redox and repair responses, suppressing DNA repair and SIRT1-mediated BBB protective pathways. Lactate is both an energy substrate and a signaling molecule, with lactylation-related mechanisms emerging in neuroinjury. Fumarate should be considered separately from fumarate esters: endogenous fumarate may influence redox and electrophile-sensitive signaling, whereas dimethyl fumarate and monomethyl fumarate are pharmacological interventions. Alpha-ketoglutarate is a TCA-cycle intermediate and cofactor for dioxygenases, but direct causal evidence linking alpha-ketoglutarate changes to BBB dysfunction in rmTBI remains limited.
Mitochondrial ROS not only act as mediators of damage but also drive the perpetuation of sterile inflammation through activation of the NLRP3 inflammasome and the NF-κB pathway. Recent studies indicate that the pulsatile release of mitochondrial ROS following repetitive mild injury serves as a critical initiating factor for microglial transition into an immune-primed state. This recurrent inflammatory stimulation, the review notes, predisposes microglia to a long-term activated phenotype that ultimately contributes to neurodegeneration. The proposed consequence is not that any individual metabolite invariably causes BBB failure, but that energy deficit, redox imbalance, and inflammatory signaling can act across the neurovascular unit, affecting endothelial cells, pericytes, astrocytes, microglia, and neurons.
Endothelial cells require ATP to maintain ion gradients, cytoskeletal organization, vesicular trafficking, and the localization of tight-junction proteins including claudin-5, occludin, and ZO-1. ATP depletion and mitochondrial ROS can therefore increase permeability and reduce recovery after injury. In rmTBI, repeated impacts may be particularly consequential when endothelial energy balance has not normalized, a temporal model that remains to be tested directly. The multicellular framework explains why BBB disruption cannot be attributed to endothelial metabolism alone and highlights the need for cell-resolved, longitudinal rmTBI experiments.
On the therapeutic front, the review identifies metabolism-directed interventions that are shifting from traditional energy supplementation toward precise regulation of the metabolic signaling network. MitoQ, a mitochondria-targeted antioxidant, has direct rmTBI evidence: C57BL/6 mice subjected to three concussive head injuries received MitoQ (5 mg/kg, intraperitoneally, twice weekly for 1 month), with attenuation of oxidative stress and chronic pathological outcomes. NAD+ restoration via precursors such as nicotinamide mononucleotide (NMN) or nicotinamide riboside (NR) effectively restores the NAD+ pool and activates SIRT1- and SIRT3-mediated mitochondrial antioxidant defense, potentially promoting BBB repair. Mdivi-1, a Drp1-mediated fission inhibitor, has shown effects on BBB disruption and cell death in experimental TBI, but this evidence is predominantly from single-TBI models and should be presented as mechanistic context rather than direct rmTBI evidence.
The review concludes that mitochondrial dysfunction and BBB impairment are shared secondary-injury processes across TBI, but in rmTBI, recurrent impacts may prolong or recouple energy stress, redox imbalance, inflammatory signaling, and neurovascular-unit dysfunction. Succinate, NAD+, lactate, fumarate, and alpha-ketoglutarate are useful candidates for testing this framework, but their causal evidence is not equivalent. Future work should report impact number and interval, injury severity, cell-specific metabolic measures, BBB endpoints, and longitudinal functional outcomes. An evidence-tiered approach is important for translation: studies should test whether timing of repeated impacts determines persistent metabolic and BBB phenotypes, and whether a given intervention restores a defined pathway rather than merely improving a broad injury score. At present, MitoQ provides direct preclinical rmTBI evidence, whereas NAD+ precursor, ethyl pyruvate, dimethyl fumarate, and mitochondrial-dynamics approaches are supported mainly by single-TBI or related experimental models and should therefore be considered candidates for appropriately powered rmTBI studies, not established treatments.
Subject of Research: Mitochondrial metabolic remodeling and blood–brain barrier dysfunction after repetitive mild traumatic brain injury
Article Title: Mitochondrial metabolic remodeling and blood–brain barrier dysfunction after repetitive mild traumatic brain injury
Article References: Gao, L., Xue, X., Deng, X., Xu, Y., & Gao, H. (2026). Mitochondrial metabolic remodeling and blood–brain barrier dysfunction after repetitive mild traumatic brain injury. Frontiers in Molecular Neuroscience, 19, Article 1852115. https://doi.org/10.3389/fnmol.2026.1852115
Image Credits: AI Generated
DOI: 10.3389/fnmol.2026.1852115
Keywords: repetitive mild traumatic brain injury, mitochondrial dysfunction, blood–brain barrier, neurovascular unit, metabolic remodeling, reactive oxygen species, NAD+ depletion, succinate, lactate, MitoQ, chronic traumatic encephalopathy, neuroinflammation
News Source: Cassandra Pierce. (October 17, 2026). Mitochondrial Metabolic Remodeling Drives Blood–Brain Barrier Dysfunction After Repetitive Mild Traumatic Brain Injury. Scienmag.