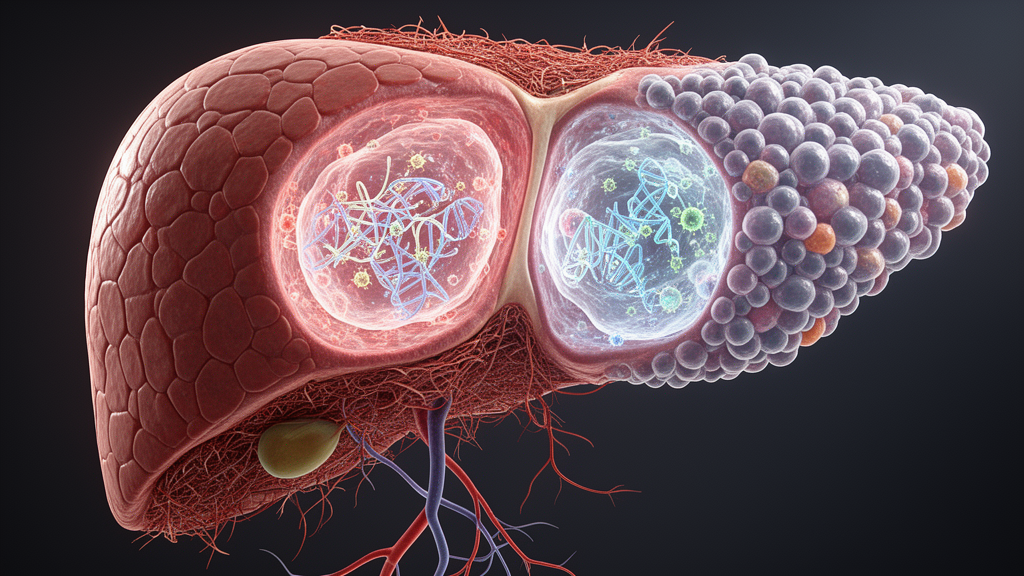
Preliminary transcriptomic profiling of liver tissue specimens has mapped differential gene expression patterns across chronic hepatitis C progression, revealing substantial transcriptional remodeling that distinguishes hepatocellular carcinoma from earlier fibrosis stages and uninfected controls.
The exploratory investigation evaluated the continuum of disease from METAVIR fibrosis stages F0 through F4 to hepatocellular carcinoma in patients with chronic hepatitis C. Hepatocellular carcinoma is the sixth most common cancer type and the third leading cause of cancer-related death worldwide, with cirrhosis associated with hepatitis B and hepatitis C virus serving as primary global risk factors accounting for approximately 80 percent of cases.
To investigate molecular mechanisms across this continuum, researchers conducted bulk RNA sequencing on liver tissue samples. The study analyzed specimens obtained from 23 chronic hepatitis C patients across METAVIR stages F0 to F4, 3 patients with hepatocellular carcinoma, and 3 uninfected liver controls, followed by differential gene expression and Gene Ontology enrichment analyses.
Tissue from hepatocellular carcinoma displayed a distinct transcriptional profile characterized by a higher number of uniquely dysregulated genes compared to uninfected controls and preceding fibrosis stages. Trajectory-oriented clustering resolved specific gene modules showing monotonic expression patterns across METAVIR stages, identifying core pathways associated with the transcriptomic shift seen in carcinoma tissue.
Coordinated expression programs included gene modules that progressively increased toward hepatocellular carcinoma, marked by proliferative and cell-cycle signals, as well as modules that decreased as fibrosis advanced, reflecting a loss of differentiated hepatocyte functions. Functional enrichment analysis further highlighted inflammatory and acute-phase responses, extracellular matrix dynamics, and lipid and cholesterol metabolism as prominent exploratory themes.
The study design utilized pooled transcriptomic libraries to optimize group-level representativeness across sample categories. The authors emphasized that these preliminary findings warrant validation in non-pooled independent cohorts, noting that further confirmation using quantitative polymerase chain reaction, protein-level assays, and external datasets including TCGA-LIHC and GEO is required to confirm candidate markers and mechanistic trajectories in chronic hepatitis C-associated hepatocarcinogenesis.
Subject of Research: Transcriptomic profiling of chronic hepatitis C progression to hepatocellular carcinoma
Article Title: The transcriptomic landscape of chronic hepatitis C progression to hepatocellular carcinoma
Article References: dos Santos, D. B., Hoyos, B. S., de Oliveira Lima, E., Silva, G. F., Galvani, A. F., Pereira, G. L., Curi, R. A., Fernandez, G. J., & Ferrasi, A. C. (2026). The transcriptomic landscape of chronic hepatitis C progression to hepatocellular carcinoma. Clinical and Experimental Medicine. https://doi.org/10.1007/s10238-026-02304-2
Image Credits: AI Generated
DOI: 10.1007/s10238-026-02304-2
Keywords: hepatocellular carcinoma, chronic hepatitis C, transcriptomics, liver fibrosis, RNA sequencing, METAVIR stages, gene expression, hepatocarcinogenesis, transcriptomic, landscape, chronic, hepatitis
News Source: Nathaniel Bowman. (November 10, 2026). Transcriptomic Shifts Trace Hepatitis C Progression to Liver Cancer. Scienmag.