A newly investigated knock-in mouse model carrying the p.R15L mutation in the endogenous CHCHD10 protein demonstrates that slowly progressive, late-onset motor neuron degeneration can occur independently of mitochondrial respiration deficits or major structural protein aggregation. Mutations in the mitochondrial protein CHCHD10 cause a spectrum of hereditary neurodegenerative disorders, and the specific p.R15L variant is associated with a late-onset familial form of amyotrophic lateral sclerosis whose pathogenic mechanisms have remained unresolved.
To elucidate the physiological consequences of this variant in vivo, researchers examined knock-in mice expressing the mutation under endogenous genetic control. The animals developed subtle, age-related physical and behavioral phenotypes as they matured, characterized by reduced overall body weight, sex-specific impairments in motor activity, and measurable anxiety-like behavior, establishing an initial clinical parallel to late-onset human neurodegenerative disease.
Unlike previously characterized knock-in models harboring different mutant CHCHD10 variants, the p.R15L mice retained normal CHCHD10 protein abundance across evaluated tissues. Biochemical and microscopic assessments showed no evidence of large protein aggregate accumulation, indicating that pathognomonic protein insolubility is not a prerequisite for phenotypic development in this genetic context.
Functional evaluations of isolated mitochondria further separated the observed pathology from classic organelle failure. In mitochondria isolated from both muscle and brain tissue of the knock-in mice, standard rates of mitochondrial respiration and levels of hydrogen peroxide emission remained completely unaltered relative to controls, demonstrating preserved respiratory chain integrity and normal reactive oxygen species production.
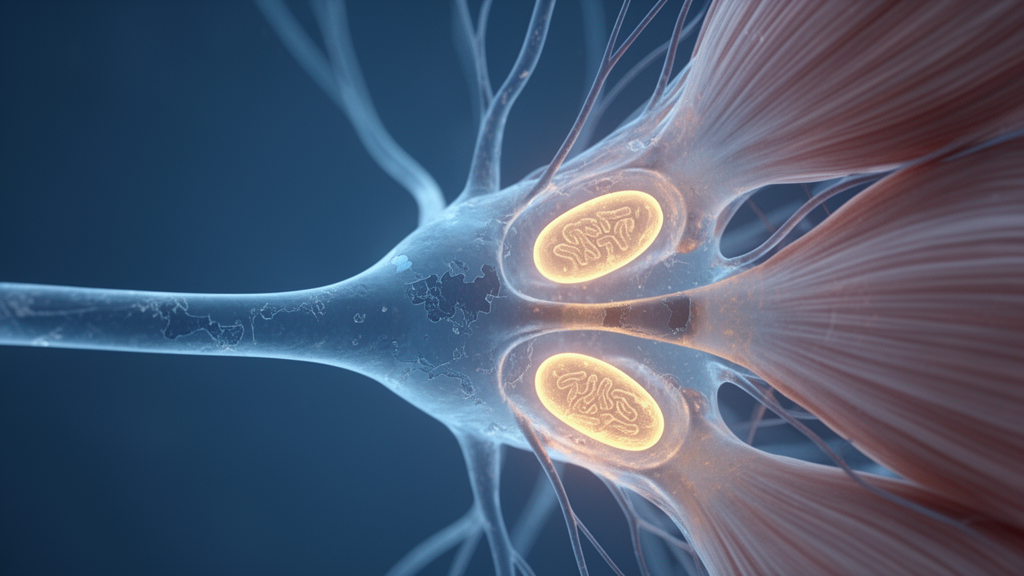
Corresponding assessments in clinical specimens confirmed that this preservation extends to human biology, as cultured fibroblasts obtained from human p.R15L mutation carriers similarly displayed normal CHCHD10 protein concentrations and intact oxidative phosphorylation function. Rather than exhibiting respiratory breakdown, histochemical evaluation of the knock-in mouse tibialis anterior muscle demonstrated increased mitochondrial enzymatic activity across a distinct subset of fibers, accompanied by RNA sequencing profiles showing transcriptional upregulation of oxidative phosphorylation genes.
Despite preserved mitochondrial organellar function, electrophysiological testing in aged mice uncovered significant neuromuscular impairment, marked by a decreased compound muscle action potential amplitude in the tibialis anterior muscle. This functional decline coincided with a moderate loss of spinal cord neurons and elevated concentrations of serum neurofilament light, confirming active neurodegeneration and supporting a model where the p.R15L mutation induces motor neuron injury primarily through a toxic gain-of-function mechanism.
The establishment of this endogenous knock-in model addresses longstanding questions regarding the pathogenic mechanisms of the p.R15L variant, which is linked to a slowly progressive, late-onset familial form of amyotrophic lateral sclerosis. Because mutations in the mitochondrial protein CHCHD10 produce a spectrum of hereditary neurodegenerative disorders, examining this mutation in the endogenous protein allows researchers to evaluate how disease manifestations unfold over time. The knock-in mice developed subtle, age-related phenotypes rather than an acute clinical deterioration, presenting with decreased body weight, sex-specific motor activity defects, and an anxiety-like behavior that partially recapitulate the progression seen in human patients.
Crucially, the experimental findings demonstrate that motor neuron impairment can arise in the complete absence of mitochondrial functional or morphological alterations. Unlike previously described mutant CHCHD10 knock-in models, the p.R15L knock-in mice exhibited normal protein levels without evidence of large protein aggregates or protein instability. This preservation was consistent across biological systems, as cultured fibroblasts obtained from human p.R15L carriers also exhibited normal CHCHD10 protein levels and unchanged oxidative phosphorylation function. Furthermore, isolated mitochondria from both muscle and brain tissues displayed unaltered mitochondrial respiration and unaltered hydrogen peroxide emission, confirming that organelle-level respiratory failure is not required for pathology to develop.
Rather than widespread organellar degradation, tissue evaluations revealed selective molecular shifts alongside measurable neuromuscular deficits. Histochemical analyses of skeletal muscle showed increases in mitochondrial enzymatic activity in a subset of tibialis anterior muscle fibers, and RNA sequencing revealed profiles indicative of increased transcription of oxidative phosphorylation genes. In aged mice, electrophysiological studies demonstrated a decreased amplitude of the compound tibialis anterior muscle action potential, commensurate with a moderate loss of spinal cord neurons and elevated serum neurofilament light levels indicative of neurodegeneration. These findings indicate that p.R15L CHCHD10 impairs mouse motor neurons, likely through a gain-of-function mechanism, offering a genetically accurate, mild in vivo model of familial amyotrophic lateral sclerosis.
Subject of Research: Motor neuron degeneration and mitochondrial function in a knock-in mouse model of familial ALS carrying the CHCHD10 p.R15L mutation
Article Title: A mouse model of CHCHD10 p.R15L familial ALS presents mild, age-related motor neuron degeneration without protein instability or mitochondrial dysfunction
Article References: Park, J., Stepanova, A., Dash, J., Southwell, N., Zhao, D., Konrad, C., Tyagi, P., Kwan, J. Y., Shneider, N. A., Mentis, G. Z., Manfredi, G., & Kawamata, H. (2026). A mouse model of CHCHD10 p.R15L familial ALS presents mild, age-related motor neuron degeneration without protein instability or mitochondrial dysfunction. Acta Neuropathologica Communications. https://doi.org/10.1186/s40478-026-02463-z
Image Credits: AI Generated
DOI: 10.1186/s40478-026-02463-z
Keywords: CHCHD10, p.R15L mutation, amyotrophic lateral sclerosis, familial ALS, motor neuron degeneration, knock-in mouse model, mitochondrial respiration, neurofilament light, tibialis anterior, compound muscle action potential, spinal cord neurons, oxidative phosphorylation, human fibroblasts, hydrogen peroxide emission, RNA sequencing, gain-of-function mechanism, mitochondrial enzymatic activity, age-related neurodegeneration
News Source: Diana Fleming. (November 14, 2026). CHCHD10 Familial ALS Mouse Model Displays Motor Degeneration Without Mitochondrial Dysfunction. Scienmag.