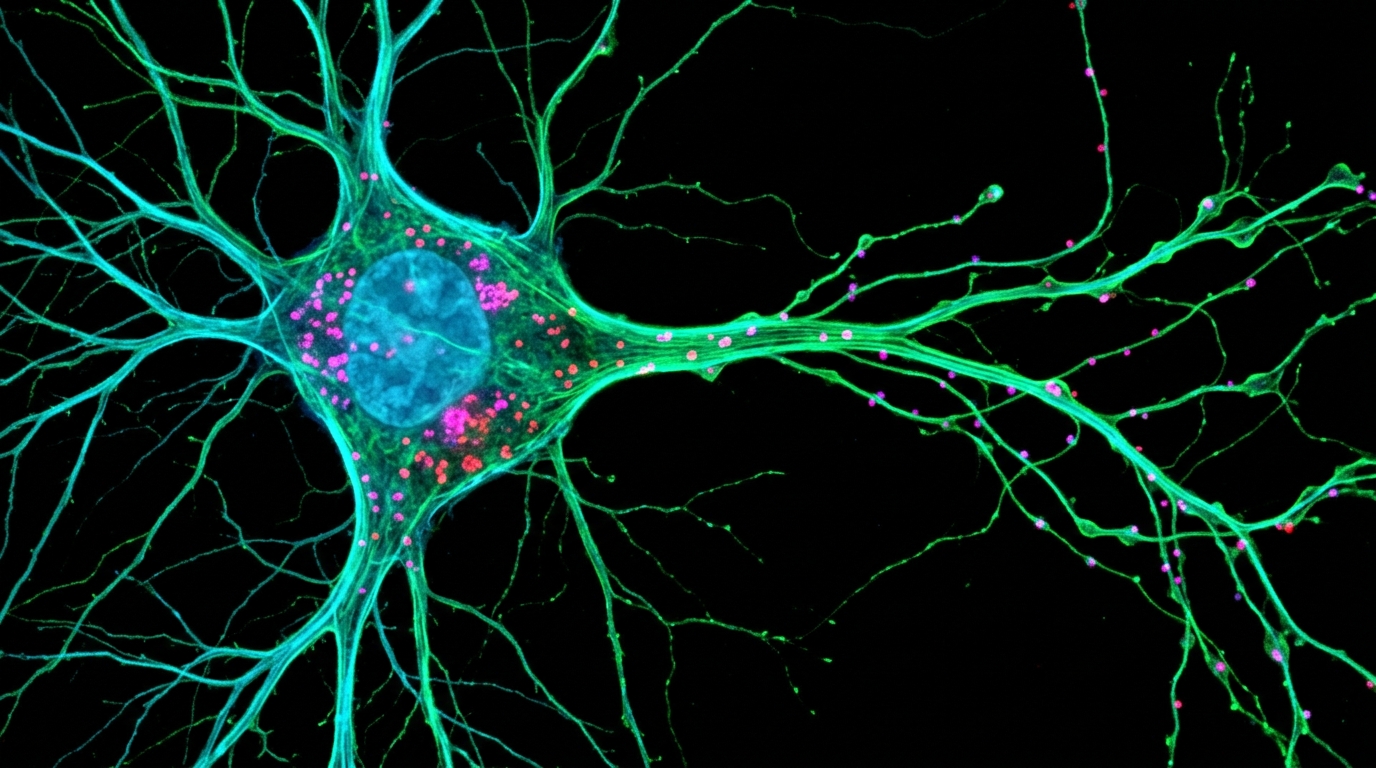
A comprehensive review published in Anesthesiology and Perioperative Science systematically details how long non-coding RNAs act as critical modulators across the mammalian nervous system to initiate, amplify, and sustain neuropathic pain. Neuropathic pain represents a chronic syndrome arising from structural lesions or disease within the somatosensory nervous system, frequently marked by persistent spontaneous discomfort, hyperalgesia, and allodynia that severely compromise patient well-being. Conventional clinical interventions, including tricyclic antidepressants, antiepileptic medications, opioids, and multidrug regimens, remain constrained by substantial adverse reactions such as nausea, dizziness, vomiting, constipation, and local erythema alongside marked inter-individual variability in analgesic efficacy. Because these painful states do not originate from isolated single receptors or linear signaling pathways, researchers emphasize that elucidating complex gene expression networks across the entire neuraxis provides an essential molecular foundation for developing targeted therapeutic strategies.
Long non-coding RNAs, designated as RNA transcripts exceeding two hundred nucleotides in length that do not encode functional proteins, have emerged from being viewed as mere transcriptional noise into recognized drivers of cellular homeostasis and gene expression. These molecules orchestrate cellular proliferation, survival, migration, and genomic integrity through direct biochemical interactions with cellular proteins, genomic DNA, and complementary RNA sequences. Across established preclinical injury models—including chronic constriction injury, sciatic nerve injury, spinal nerve ligation, and diabetic neuropathic pain models—high-throughput RNA sequencing and microarray profiling have revealed extensive dysregulation of long non-coding RNA expression. When evaluated alongside weighted gene co-expression network analysis and competing endogenous RNA network modeling, these transcriptomic alterations coordinate aberrant neuronal excitability, the activation of non-neuronal glial cells, and neuroinflammatory signaling cascades that drive sensory pathology.
At the most peripheral level of sensory detection, damage to peripheral nerve tissue triggers dramatic transcriptional remodeling within local nerve fibers and their surrounding microenvironment. Microarray investigations in sciatic nerve injury models show that hundreds of long non-coding RNAs undergo significant upregulation or downregulation, directly influencing repair-associated biological processes including Schwann cell proliferation, axonal regeneration, and distal ion channel distribution. In spinal nerve ligation paradigms, long non-coding RNA H19 exhibits marked induction across peripheral Schwann cells, dorsal root ganglia, and astrocytes, while signaling cascades such as SIX1/HULC exacerbate hydrogen-peroxide-induced Schwann cell injury. Although peripheral nerve injury reliably elevates potent inflammatory cytokines such as tumor necrosis factor-alpha and interleukin-1-beta, authors note that technical limitations in isolating peripheral nociceptor terminals have historically hindered concurrent profiling of terminal non-coding transcripts and local cytokine cascades, leaving precise peripheral mechanisms an active subject of ongoing investigation.
Within the dorsal root ganglia, which serve as the primary sensory relay stations transmitting incoming peripheral signals toward the central nervous system, extensive transcriptomic reconfiguration directly drives sensory neuron hyperexcitability. Rodent models of sciatic nerve injury exhibit over one hundred differentially expressed long non-coding RNAs that coordinate the functional remodeling of membrane receptors and ion channels. Specifically, upregulated transcripts such as MRAK009713 and NONRATT021972 elevate the expression and functional activity of purinergic P2X3 and P2X7 receptors, stimulating downstream extracellular signal-regulated kinase pathways and triggering proinflammatory mediator release. Concurrently, lncRNA BC168687 coordinates aberrant activation across both sensory neurons and satellite glial cells under diabetic conditions through P2X7 and transient receptor potential vanilloid 1 modulation. Electrophysiological stability is further disrupted as Kcna2 antisense RNA suppresses voltage-gated potassium channel Kv1.2 expression, while sensory neuron-specific non-coding transcripts fail to properly bind KCNN1 promoters, reducing repolarizing currents and allowing sustained sodium-channel-driven firing.
As primary afferent nociceptive inputs enter the spinal cord dorsal horn, dysregulated long non-coding RNAs govern the amplification of pain signaling and the progression of central sensitization. High-throughput sequencing of spinal segments L3 through L5 in chronic constriction injury models demonstrates broad alterations in non-coding transcripts heavily enriched in cytokine production and intercellular communication networks. For example, spinal upregulation of lncRNA PCAT19 relieves microRNA-378a-3p repression as a competing endogenous RNA, freeing KDM3A to demethylate the brain-derived neurotrophic factor promoter and aggressively stimulating microglial neuroinflammation. In parallel, non-cell-cycle functions of cyclin-dependent kinases contribute to spinal hypersensitivity; while cyclin-dependent kinase 5 phosphorylates TRPV1 to drive inflammatory thermal hypersensitivity, spinal nerve ligation induces lncRNA PKIA-AS1 to hypomethylate the CDK6 promoter, elevating interleukin-1-beta, interleukin-6, interleukin-12, and tumor necrosis factor-alpha to reinforce chronic spinal transmission.
Ascending pain pathways subsequently project signals into higher-order cerebral structures, where long non-coding RNAs actively participate in the emotional, affective, and cognitive comorbidity frequently accompanying persistent physical discomfort. Causal inference investigations have demonstrated positive causal associations between chronic pain across multiple anatomical regions and seven major psychiatric conditions, including depression, panic disorder, generalized anxiety, insomnia, attention-deficit/hyperactivity disorder, post-traumatic stress disorder, and bipolar disorder. Within the anterior cingulate cortex and hippocampus, which form a central emotional processing axis, lncRNA CCAT1 downregulation impairs miR-155/SGK3 regulatory signaling, whereas diabetic neuropathic pain models reveal hippocampal upregulation of MSTRG.81401 driving astrocyte P2X7-mediated NLRP3/caspase-1 pyroptotic neuroinflammation. Within the central amygdala, reduced lncRNA-84277 expression relieves repression on miR-128-3p to suppress protective SIRT1 enzymes, exacerbating depressive phenotypes that can be reversed by experimental viral restoration.
Beyond anatomical spatial distribution, long non-coding RNAs display distinct temporal dynamics as neuropathic conditions progress from acute post-injury phases into established, chronic states. Preclinical evidence indicates that transcripts such as H19 and NEAT1 exhibit pronounced upregulation during early acute stages to drive initial peripheral sensitization, whereas distinct regulators including KCNA2-AS and MEG3 maintain elevated expression over prolonged periods to sustain persistent glial activation and long-term neuroinflammation. However, scientific evaluation of these temporal shifts remains hindered by methodological challenges, as dynamic profiling across multiple disease stages requires large animal cohorts and repeated harvesting of delicate neural structures like the spinal cord and dorsal root ganglia from individual subjects. Low baseline transcript abundance, high cell-type specificity, variable pain behaviors, and fluctuating inflammatory microenvironments create substantial data noise that current longitudinal experimental designs are working to resolve.
These detailed mechanistic insights have fostered diverse therapeutic strategies designed to manipulate pain-associated long non-coding RNAs through targeted molecular intervention. Antisense oligonucleotides and structurally optimized GapmeRs utilize complementary base pairing to initiate RNase H-mediated RNA degradation, showing preclinical efficacy by significantly reversing mechanical and thermal hypersensitivity following intrathecal or subcutaneous delivery in nerve injury models. Complementary RNA interference platforms using short interfering RNAs or viral-vector-delivered short hairpin RNAs successfully depress neuroinflammatory cytokine release and restore pain thresholds, exemplified by SNHG12 silencing in injured dorsal root ganglia. Genomic editing via CRISPR-Cas9 offers irreversible targeted sequence disruption, while replacement therapies using neurotropic viral vectors demonstrate that restoring deficient protective transcripts like spinal lncRNA FTX can suppress spinal cord inflammation, complemented by synthetic small molecules that influence broader RNA stability networks.
Despite compelling preclinical outcomes in animal models, the clinical translation of long non-coding RNA interventions faces substantial biological and pharmacological hurdles. Long non-coding RNAs exhibit remarkably poor evolutionary conservation across species; human transcripts frequently contain abundant Alu transposable elements that are virtually absent in rodent sequences, meaning many targets identified in preclinical models lack functional human orthologs. An important exception involves transcriptionally ultra-conserved regions, such as uc.153, which maintain strict multi-species conservation and regulate microRNA maturation during neuropathic progression. Furthermore, non-coding transcripts display extreme tissue, cellular, and subcellular specificity, where nuclear molecules direct transcriptional complexes while cytoplasmic counterparts modulate translation, creating risks of bidirectional or tissue-opposing effects if targeted systemically.
Overcoming these translational barriers will require innovative delivery platforms and clinically relevant experimental models capable of navigating the mammalian nervous system safely. Because unmodified therapeutic nucleic acids cannot cross the intact blood-brain barrier, preclinical models rely heavily on invasive intrathecal or intracerebroventricular injections that pose significant procedural risks of infection, localized inflammation, and poor compliance in chronic human populations. Future translational research is prioritizing engineered delivery vectors such as natural exosomes and surface-modified nanoparticles designed to cross the blood-brain barrier non-invasively, combined with human induced pluripotent stem cell sensory neurons and organoids for human target validation. Paired with artificial intelligence tools capable of predicting secondary RNA structures and designing precise oligonucleotide sequences, mapping non-coding RNA networks offers a promising path toward personalized therapies for chronic neuropathic pain.
Subject of Research: Molecular mechanisms and therapeutic potential of long non-coding RNAs in neuropathic pain
Article Title: Long non-coding RNAs as key modulators in neuropathic pain: mechanistic pathways and novel therapeutic opportunities
Article References: Guo, L., Sun, G., Li, L., & Han, G. (2026). Long non-coding RNAs as key modulators in neuropathic pain: mechanistic pathways and novel therapeutic opportunities. Anesthesiology and Perioperative Science, 4(1), Article 23. https://doi.org/10.1007/s44254-026-00172-3
Image Credits: AI Generated
DOI: 10.1007/s44254-026-00172-3
Keywords: neuropathic pain, long non-coding RNA, neuroinflammation, dorsal root ganglia, spinal dorsal horn, central sensitization, antisense oligonucleotides, RNA interference, gene regulation, translational medicine, Long, non-coding
News Source: Juliet Wilcox. (November 9, 2026). Researchers Map How Long Non-Coding RNAs Modulate Neuropathic Pain Pathways. Scienmag.