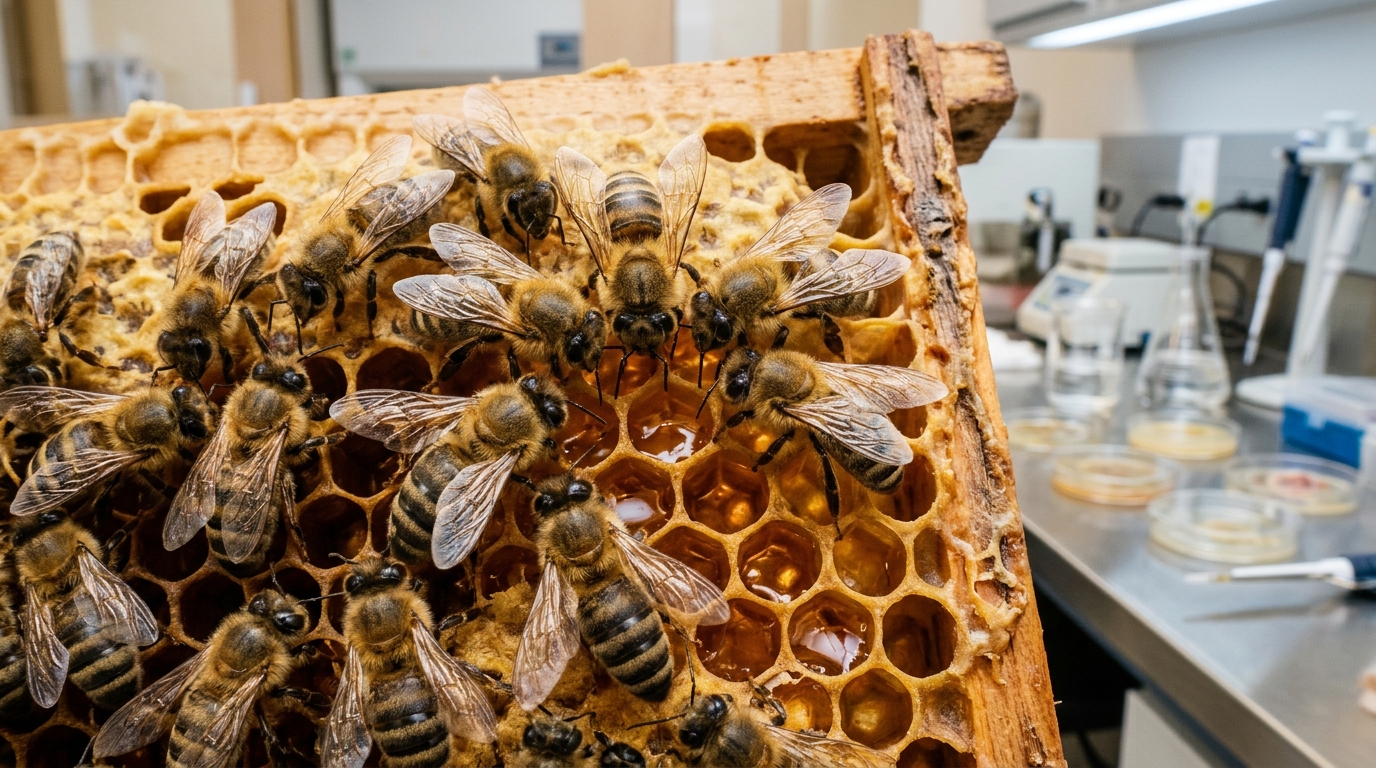
The honey bee colony behaves as a single biological unit, a superorganism whose members share labor, food and a collective physiology. Yet the microbial communities that live within this superorganism have been described almost entirely from one tissue: the hindgut of adult workers. A new study argues that this narrow view has been reinforced by technical choices that most researchers never see, from which stretch of the 16S rRNA gene is sequenced to how sequencing errors are filtered and which reference database assigns species names. The result, published in Microbial Ecology, is a sweeping reanalysis showing that conclusions about honey bee microbiota depend strongly on method, not just biology.
Duan C. Copeland, Brendon M. Mott, Charles J. Mason and Kirk E. Anderson reprocessed 4,586 short-read Illumina libraries drawn from 37 BioProjects, then added 345 full-length PacBio libraries covering the entire V1-V9 region of the 16S rRNA gene, generated from dissected worker and queen gut tissues to serve as a reference. By running every library through a single set of filtering and reporting thresholds with one species-level reference database, the team removed the confounding that normally prevents direct comparison across studies and niches. Each library was processed twice, once with read denoising and once without, to isolate the effect of that single bioinformatic decision.
The two processing routes agreed on which genera were present, but they diverged sharply on finer detail. Denoised reads produced far fewer amplicon sequence variants per library and significantly lower alpha diversity, meaning that a routine pipeline choice visibly reshaped the apparent richness of the community. Species-level resolution proved even more dependent on the sequenced region: full-length reads named nearly every major genus down to species, while some short variable regions named almost none. Studies that rely on a single short region may therefore be systematically blind to species identity, regardless of how carefully the data are analyzed.
Looking across niches rather than at the worker hindgut alone, the authors found that the superorganismal microbiota is composed of both niche specialists and generalists shared across multiple body regions and colony environments. They also identified at least three abundant amplicon sequence variants that lack any species-level description, suggesting that even the most intensively sampled insect microbiota still contains unnamed organisms. The full-length reference libraries provide a template for naming them in future work.
The study also documents a persistent and underappreciated problem: contaminant reads. Because next-generation sequencing is sensitive enough to detect microbes in niches with very low bacterial biomass, contamination scaled inversely with bacterial load. Contaminants dominated drone reproductive tissue, larvae, mouthparts, hive materials, control or lab-reared worker guts and stored food. In such low-biomass samples, the true resident community can be drowned out by sequences introduced during sample handling or sequencing itself, and the lower the bacterial load, the worse the distortion becomes.
The practical implication is that any claim about honey bee microbiota, or about how the microbiota integrates with host traits such as age, caste or behavior, must be read alongside three technical variables: the primer region used, how sequencing error was handled, and the level of high-throughput sequencing contamination. Two studies of the same tissue can reach different conclusions for reasons that have nothing to do with the bees themselves. The authors frame their work as a step toward a genuinely superorganismal perspective, one in which the colony’s microbial identity is mapped across all of its niches with methods that can be compared directly.
This reanalysis does not overturn previous findings, but it supplies a standardized reference against which future studies can be calibrated, and it warns that the superorganism’s microbial portrait is still being drawn.
Subject of Research: Taxonomic resolution of the honey bee superorganismal microbiota across niches and 16S rRNA gene regions
Article Title: Towards a Superorganismal Perspective of Honey Bee Microbiota: Taxonomic Resolution by Niche and 16S rRNA Gene Variable Region
Article References: Copeland, D. C., Mott, B. M., Mason, C. J., & Anderson, K. E. (2026). Towards a Superorganismal Perspective of Honey Bee Microbiota: Taxonomic Resolution by Niche and 16S rRNA Gene Variable Region. Microbial Ecology. https://doi.org/10.1007/s00248-026-02923-y
Image Credits: AI Generated
DOI: 10.1007/s00248-026-02923-y
Keywords: honey bee, microbiota, superorganism, 16S rRNA gene, amplicon sequencing, denoising, contamination, niche specialists, alpha diversity, PacBio, species resolution, Towards
News Source: Morgan Morrow. (November 6, 2026). Honey Bee Colonies Are Superorganisms, and Their Microbiota Depends on How You Sequence It. Scienmag.