Sodium-ion batteries are racing to become the affordable backbone of grid-scale energy storage, but their success hinges on a microscopic drama that unfolds at the electrode-electrolyte interface. Now, a team of researchers has used density functional theory (DFT) calculations to watch that drama play out atom by atom, revealing how five common electrolyte anions interact with molybdenum disulfide (MoS2) surfaces as the battery charges and discharges. The findings, published in Frontiers in Batteries and Electrochemistry, offer some of the most detailed atomistic insight yet into how the protective solid electrolyte interphase (SEI) might form on one of the most promising next-generation anode materials.
The stakes are considerable. Lithium-ion batteries dominate the market, but the high cost of lithium salts and the limited availability of terrestrial lithium resources have pushed scientists toward alternatives. Sodium, abundant in the Earth’s crust and inexpensive to extract, is the obvious candidate. Yet sodium ions are bulkier than lithium ions, with an ionic radius of 1.02 angstroms compared to 0.76 angstroms, and they carry a higher reduction potential and slower reaction kinetics. That makes finding suitable electrode materials harder. MoS2, a layered two-dimensional transition metal dichalcogenide with a theoretical capacity of roughly 670 mAh/g, has emerged as a leading contender thanks to its natural abundance, straightforward synthesis, and generous interlayer spacing that eases ion intercalation.
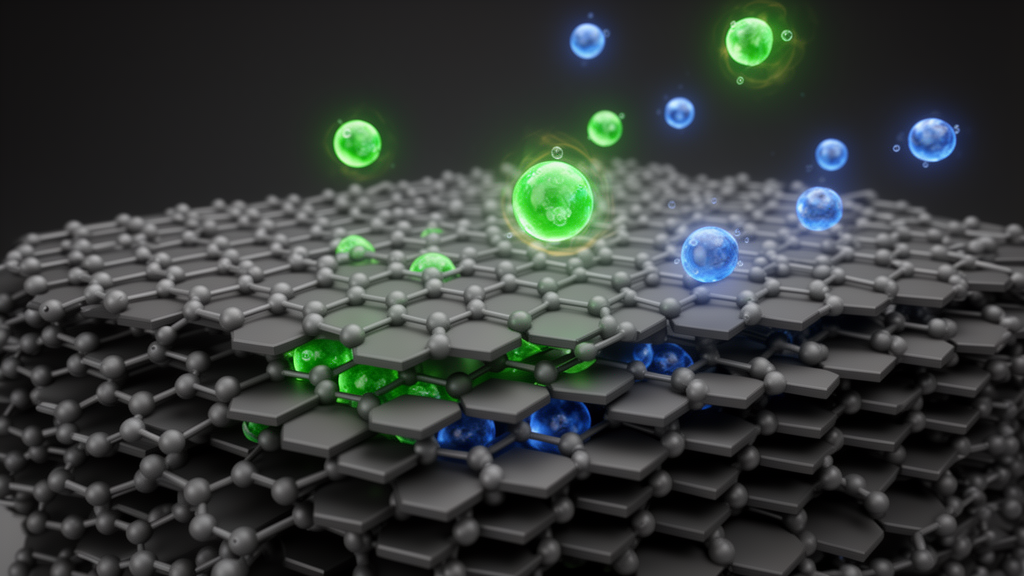
The research team, led by Luis A. Selis of Universidad Nacional de Ingeniería in Peru, Utkarsh Vijay, and Diego E. Galvez-Aranda of Texas A&M University, focused on the stable 2H phase of MoS2, the polymorph most widely studied for battery applications. Using the Quantum ESPRESSO software package with the Perdew-Burke-Ernzerhof exchange-correlation functional, they built periodic simulation cells containing a 48-atom MoS2 slab and a single electrolyte anion. The five anions examined were hexafluorophosphate (PF6-), tetrafluoroborate (BF4-), perchlorate (ClO4-), bis(fluorosulfonyl)imide (FSI-), and bis(trifluoromethanesulfonyl)imide (TFSI-), all staples of real battery electrolytes.
Crucially, the team did not model the surface at a single static state. Instead, they simulated three degrees of sodiation, corresponding to 0 percent, 50 percent, and 90 percent sodium content in the electrode, capturing how the interface evolves as the battery fills with sodium. For each anion and each sodium concentration, multiple molecular orientations were tested, and the researchers evaluated interaction energies, charge redistribution, the electron localization function, work function changes, and electronic structure modifications. The models deliberately exclude explicit solvent molecules and reactive decomposition pathways, but the authors note that similar simplified interface models have been widely used in prior DFT studies to probe early interfacial processes relevant to SEI evolution.
The results paint strikingly different portraits for each anion. Tetrafluoroborate, BF4-, behaves like a committed guest that changes its mind. At 0 percent sodium, it binds strongly to the MoS2 surface, with charge density difference maps showing pronounced electron accumulation at a fluorine atom contacting the surface, suggesting partial bond formation and even a possible weakening of the B-F bond. But as sodiation increases, the interaction collapses: at 50 percent sodium the interaction energy turns slightly positive, and at 90 percent it reaches +0.33 eV, an outright unfavorable configuration. The work function difference progressively increases, indicating reduced electronic perturbation of the anode as sodium accumulates.
Perchlorate, ClO4-, tells a subtler story. Its interaction energy starts strongly negative at -2.55 eV on the pristine surface but weakens to -1.34 eV and -1.09 eV at 50 and 90 percent sodium. What makes this case intriguing is that the total charge transfer barely changes across the entire range, remaining between 0.10 and 0.16 electrons. The weakening, the researchers found, is instead driven by a spatial redistribution of electrons: the electron localization function reveals growing localization in regions associated with intercalated sodium atoms rather than the MoS2 surface itself. In other words, the anion keeps transferring charge, but the sodium steals the electronic spotlight.
Hexafluorophosphate, PF6-, is the outright failure of the group. It never forms a stable interface configuration on MoS2 at any sodium concentration. At 0 percent sodium it sits tilted against the surface with no preferential interaction, and as sodium content rises it reorients toward a vertical configuration while sodium atoms migrate toward its fluorine atoms. The interaction energies turn positive, reaching +0.44 eV at 90 percent sodium, and the anion effectively abandons the surface in favor of bonding with intercalated sodium. This finding matters because PF6- is among the most common salts in commercial electrolytes, and its inability to anchor to MoS2 could shape how the SEI nucleates.
The large, flexible anions TFSI- and FSI- display the most complex behavior, governed by molecular orientation. For TFSI-, only oxygen-coordinated configurations proved favorable at low sodium, with the O-attack orientation yielding -0.49 eV while fluorine-dominated orientations remained energetically unfavorable at positive energies. Molecular electrostatic potential calculations explain why: the regions around oxygen atoms are consistently more negative than those near fluorine atoms, making oxygen the preferred contact point with the positively charged molybdenum and sodium sites. FSI- showed something even more dramatic. In two of its three tested orientations, at 90 percent sodium the S-F bond within the anion spontaneously dissociated, with the freed fluorine atom forming a stable bond with a surface sodium atom. The interface energies plunged to -8.64 eV and -9.06 eV, and charge transfer surged to -1.29 and -1.54 electrons, marking a transition from a molecular interface to a dissociative one.
That spontaneous FSI- decomposition is arguably the paper’s most consequential discovery. The authors conclude that highly sodiated surface regions dramatically lower the energetic barrier for FSI- decomposition, identifying the Na-F interaction as the primary driving force for initial electrolyte degradation. Since electrolyte decomposition is precisely what builds the SEI, the thin passivating layer that protects an anode from further parasitic reactions, this suggests that the state of charge itself dictates when and how the interphase begins to form. A general trend also emerged across all five anions: increasing sodium concentration weakens the direct anion-surface interaction while promoting sodium-mediated interactions, a shift the authors link to the changing electrostatic potential of the electrode, which concentrates around both molybdenum and intercalated sodium ions as sodiation deepens.
The work is candid about its limits. Static DFT calculations cannot capture the dynamic behavior of a complete electrolyte environment with salt and solvent molecules, and the authors position their approach as an initial screening framework ahead of more computationally demanding ab initio molecular dynamics simulations. Still, the practical payoff is real: the calculated interface energies, preferred orientations, charge redistribution patterns, and work-function shifts provide atomistic descriptors that could guide the selection of electrolyte salts for MoS2-based sodium-ion batteries before any experimental validation. Smaller, symmetric anions behave predictably, while larger, flexible ones produce complex, orientation-dependent interfaces, and electronegative oxygen atoms emerge as key stabilizers. As sodium-ion batteries push toward commercialization for renewable energy storage, understanding these first atomic moments at the electrode surface may prove just as important as the electrodes themselves.
Subject of Research: First-principles simulation of electrolyte anion stability and orientation on MoS2 surfaces for solid electrolyte interphase formation in sodium-ion batteries
Article Title: First-principles study of electrolyte components stability and orientation on MoS2 surfaces: implications for SEI formation in sodium-ion batteries
Article References: Selis, L. A., Vijay, U., & Galvez-Aranda, D. E. (2026). First-principles study of electrolyte components stability and orientation on MoS2 surfaces: implications for SEI formation in sodium-ion batteries. Frontiers in Batteries and Electrochemistry, 5, Article 1845442. https://doi.org/10.3389/fbael.2026.1845442
Image Credits: AI Generated
DOI: 10.3389/fbael.2026.1845442
Keywords: sodium-ion batteries, MoS2, solid electrolyte interphase, density functional theory, electrolyte anions, SEI formation, anode materials, charge transfer, sodiation, work function, electrode-electrolyte interface, FSI decomposition
News Source: Faith Mcneil. (October 16, 2026). Atomic-Scale Simulations Reveal How Electrolyte Anions Behave on MoS2 Battery Anodes. Scienmag.