Epigenetic modifications across the mammalian genome play decisive roles in shaping neural identity and governing cellular specialization, yet standard analytical approaches have faced severe chemical limitations in distinguishing structurally related nucleotide marks. Cytosine bases can undergo methylation at the fifth carbon position to generate 5-methylcytosine, an alteration historically recognized for establishing transcriptional repression, genomic imprinting, and transposable element silencing. Through iterative oxidation driven by Ten-Eleven Translocation enzymes, 5-methylcytosine converts into 5-hydroxymethylcytosine, an epigenetic modification that prominently accumulates within neuronal gene bodies and active regulatory elements. Conventional analytical workflows rely upon bisulfite treatment, a chemical conversion process that converts unmodified cytosines into uracils while leaving both 5-methylcytosine and 5-hydroxymethylcytosine completely intact. Because bisulfite chemistry treats both modifications identically, standard assays generate an indistinguishable composite signal that obscures base-specific regulatory transitions. To overcome these constraints, researchers engineered Joint-Cabernet, a bisulfite-free single-cell sequencing platform capable of simultaneously capturing the cellular transcriptome alongside distinct, high-resolution measurements of 5-methylcytosine and 5-hydroxymethylcytosine across tens of thousands of individual nuclei isolated from the adult mouse brain.
Developing an assay capable of concurrent multi-omic interrogation without splitting delicate cellular lysates required fundamental redesigns of molecular barcoding and enzymatic chemistry. Prior bisulfite-free techniques suffered from substantial limitations, including degradative DNA loss, low chemical conversion efficiencies falling below ninety percent, or strict dependencies on restriction endonuclease recognition motifs that undersampled dense CpG islands. The foundational Cabernet protocol utilized Tn5 transposase fragmentation coupled with protective enzymatic cascades, but its direct transposon indexing strategy exhibited poor insertion uniformity across single cells. In standard Tn5 tagmentation workflows, up to fifty percent of generated genomic fragments are discarded because viable flow-cell sequencing libraries require the stochastic incorporation of two distinct adapters on opposite ends of each DNA strand. To bypass this thermodynamic inefficiency, Joint-Cabernet implements a universal symmetrical strand transposome armed with a uracil-containing linker and a cytosine-depleted reverse adapter. During initial gap-fill synthesis, the incorporated uracil residues arrest polymerase activity, enabling subsequent template-guided extension with custom locked nucleic acid oligonucleotides containing inline cellular barcodes, ensuring uniform genome recovery without generating artifactual methylation signals during amplification.
To capture transcriptomic data concurrently from identical single nuclei without physical separation of lysate, Joint-Cabernet integrates an in situ reverse transcription step directly prior to genomic tagmentation. Intact nuclei isolated from brain tissues undergo reverse transcription driven by a specialized poly-T primer engineered with a distinct reverse adapter showing zero sequence homology with genomic DNA adapters, permitting targeted amplification of both molecular species from the same reaction vessel. Furthermore, the reaction replaces natural deoxycytidine triphosphate with 5-hydroxymethyl-dCTP, forcing reverse transcriptase to synthesize fully hydroxymethylated complementary DNA strands. Because endogenous non-CpG methylation occurs at exceptionally low frequencies throughout native mammalian genomes, this pervasive synthetic hydroxymethylation functions as an unambiguous molecular identifier. During bioinformatic processing, reads exhibiting non-CpG modification frequencies of seventy percent or greater are classified as RNA-derived sequences, while reads displaying non-CpG methylation below fifty percent are classified as DNA-derived fragments, eliminating cross-contamination between genomic and transcriptomic sequencing libraries.
Accurate quantification of both cytosine modifications relies upon a two-channel enzymatic bifurcation following single-nucleus pooling. Barcoded genomic material from ninety-six-well plates is divided evenly into two separate analytical processing channels. In the first channel, combined treatment with TET2 dioxygenase and beta-glucosyltransferase protects both 5-methylcytosine and 5-hydroxymethylcytosine from downstream enzymatic alteration. In the second channel, beta-glucosyltransferase is administered alone, transferring a glucose moiety exclusively to 5-hydroxymethylcytosine while leaving 5-methylcytosine completely unprotected. Both channels then undergo deamination via APOBEC3A deaminase, an enzyme that selectively converts all unprotected cytosine residues into uracil. Because the barcoded primers incorporate modified nucleotides, synthetic barcodes remain fully protected against deamination. Quality control benchmarks demonstrated 99.5 percent accuracy for 5-hydroxymethylcytosine detection using control spike-ins and 98.3 percent accuracy for combined 5-methylcytosine and 5-hydroxymethylcytosine quantification. True 5-methylcytosine concentrations are subsequently calculated at single-cell resolution by bioinformatically subtracting the measured 5-hydroxymethylcytosine signal of the second channel from the combined modified cytosine signal obtained in the first channel.
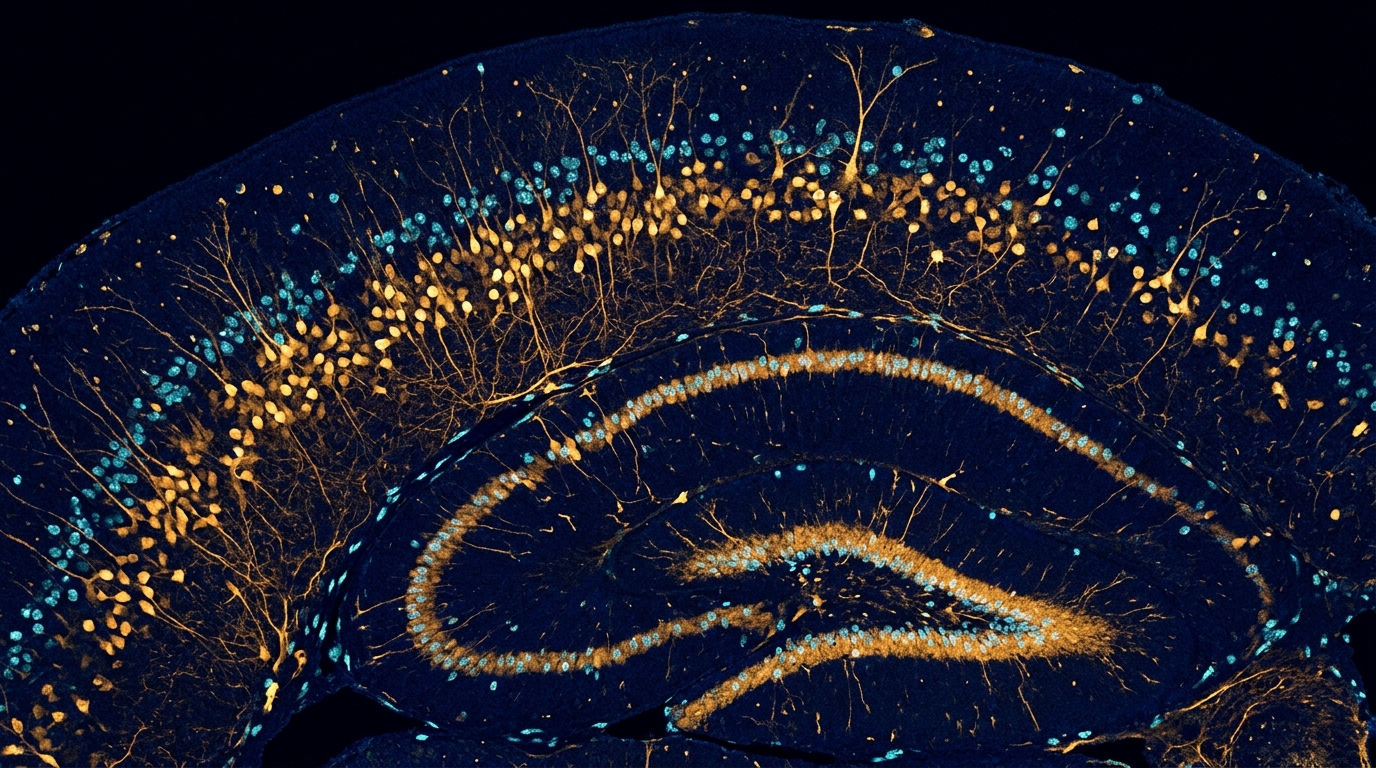
Applying Joint-Cabernet to tissue dissected from the cortex, hippocampus, striatum, and olfactory bulb of adult mice aged six to ten weeks yielded 47,712 profiled nuclei. Nuclei were labeled with anti-NeuN antibodies and isolated via fluorescence-activated cell sorting, resulting in 44,689 RNA-qualified nuclei and 43,075 high-quality cells that passed rigorous epigenomic filtering standards requiring over 98 percent accuracy and less than one percent unmodified background. Sequencing achieved an average of 8.2 million raw reads per cell for hydroxymethylation and 8.3 million raw reads per cell for combined modifications, each aligning to cover approximately 1.4 million CpG sites. Transcriptomic profiling detected an average of 2,741 expressed genes per neuron and 1,352 genes per non-neuronal cell. Projecting transcriptomic vectors onto reference single-cell atlases through robust principal component analysis identified 13 major cell classes and 31 distinct subclasses, including diverse excitatory projection neurons and inhibitory interneurons corresponding faithfully to anatomical origin tissues.
Evaluating global epigenetic marks revealed profound divergence across cellular lineages that had remained entirely invisible under conventional bisulfite profiling. While the composite measurement of 5-methylcytosine plus 5-hydroxymethylcytosine remained largely flat across diverse neuronal and glial cell types, true base-specific levels varied substantially. Fully differentiated, postmitotic mature neurons exhibited markedly elevated global 5-hydroxymethylcytosine levels compared to actively dividing non-neuronal cells. In non-dividing mature neurons, the absence of DNA replication prevents replication-dependent dilution of modified bases, allowing TET enzymes to chronically deposit and preserve hydroxymethylation across the cellular lifespan. Subtype comparisons demonstrated that somatostatin-positive GABAergic interneurons contained the highest global concentrations of 5-hydroxymethylcytosine. In contrast, immature and proliferative neuronal populations residing within the dentate gyrus and olfactory bulb exhibited depleted levels of hydroxymethylation, reflecting continuous dilution driven by ongoing adult neurogenesis. Non-CpG modifications mirrored true non-CpG methylation almost entirely, demonstrating that oxidative modification by TET enzymes occurs predominantly at canonical CpG motifs.
By integrating matched transcriptomic vectors directly with gene body epigenomic measurements, the investigators resolved the precise relationship between cytosine modifications and transcriptional activity across the genome. When genes were stratified into non-expressed, low-expressed, and high-expressed cohorts, active transcription universally correlated with severe depletion of gene body 5-methylcytosine accompanied by a massive enrichment of 5-hydroxymethylcytosine. Because the simultaneous reduction of methylcytosine and accumulation of hydroxymethylcytosine balanced each other, the composite bisulfite-like signal across active gene bodies remained static. At transcription start sites, absolute quantities of both modifications were depleted, consistent with chromatin opening required for transcription factor assembly, although the relative ratio of 5-hydroxymethylcytosine to total modifications spiked significantly. For example, the layer-six corticothalamic marker gene Cdh18 exhibited dramatic gene body hydroxymethylation in its expressing cell type, whereas its combined bisulfite metric remained completely unchanging across cortical layers, confirming that conventional assays fail to detect cell-type-specific activation signatures.
Topological mapping against three-dimensional chromatin architecture revealed that cytosine hydroxymethylation tracks open, transcriptionally active genomic domains. Analyzing Hi-C chromosomal contact matrices partitioned into active A compartments and densely packed, repressed B compartments demonstrated that A compartments are enriched for 5-hydroxymethylcytosine and depleted of 5-methylcytosine. Across one-hundred-kilobase genomic bins, positive compartment score shifts reflecting structural transitions toward open chromatin correlated positively with 5-hydroxymethylcytosine and negatively with 5-methylcytosine, an opposing relationship obscured by composite signals. Examination of cis-regulatory elements identified through ATAC-seq revealed that open chromatin centers exhibit localized cratering of all cytosine modifications, likely reflecting rapid enzymatic progression to unmodified cytosine within physically accessible nucleosome-depleted cores. In contrast, the immediate flanking boundaries of these regulatory elements displayed intense epigenetic editing, characterized by heavy 5-hydroxymethylcytosine accumulation and profound 5-methylcytosine depletion, aligning with active enhancer histone marks such as H3K4me1 and H3K27ac.
Evaluating thousands of genes displaying positive correlations with hydroxymethylation uncovered two distinct, functionally specialized regulatory mechanisms governing transcription in neuronal lineages. Among 3,337 genes exhibiting strong positive associations with 5-hydroxymethylcytosine, 97.5 percent showed strict negative correlations with 5-methylcytosine. Group 1 comprised 2,376 dynamic genes that undergo a progressive multi-step transition: 5-methylcytosine is first converted into 5-hydroxymethylcytosine during moderate gene activation, followed by an extended transition to fully unmodified cytosines during maximal expression. These genes possess longer genomic sequences with a median length of 109.7 kilobases, contain a median of 1,126 CpG sites, exhibit high expression variability across cell types, and are heavily enriched for neurodevelopmental and synaptic processes. In contrast, Group 2 comprised 840 genes where transcriptional activation is linked solely to the stable maintenance of 5-hydroxymethylcytosine without subsequent conversion to unmodified cytosine. These genes are significantly shorter with a median length of 61.5 kilobases, contain fewer CpG sites, show lower transcriptional variability, and govern essential metabolic and cellular housekeeping functions, demonstrating that stable hydroxymethylation acts as a persistent regulatory mark.
Analysis of cell-type-specific differentially hydroxymethylated regions identified 200,438 genomic sites across neuronal subclasses, with 153,580 showing hyper-hydroxymethylation in at least one subtype. Linking distal intergenic regions to target promoters using chromosome conformation capture identified 4,105 expression-associated interaction anchors. For example, the neurodevelopmental gene Vwc2 displayed elevated expression in parvalbumin-expressing interneurons alongside an enriched interaction with an upstream distal anchor located 255 kilobases away that exhibited pronounced 5-hydroxymethylcytosine accumulation and 5-methylcytosine depletion. To restore cytoarchitectural context lost during tissue dissociation, the researchers computationally projected Joint-Cabernet profiles onto spatial coordinates from high-resolution MERFISH reference maps. This location imputation uncovered geographical epigenetic gradients across cortical and hippocampal structures, including coordinated expression and gene body hydroxymethylation gradients for glutamate receptor Grik3 and axon guidance receptor Ephb1, establishing that cytosine modifications are organized in precise regional patterns across complex tissues.
Despite providing unprecedented multi-omic resolution, the authors highlighted important methodological boundaries inherent to the current Joint-Cabernet workflow. Because the platform relies on physical splitting into two enzymatic channels, 5-methylcytosine and 5-hydroxymethylcytosine cannot be co-detected on the exact same individual DNA molecules, necessitating binning across genomic windows to mitigate single-cell coverage sparsity. Furthermore, the plate-based tagmentation protocol demands greater manual processing than droplet-based single-cell platforms, while integrated in situ transcriptomic profiling yields lower gene detection sensitivity than dedicated single-cell RNA sequencing assays. Additionally, utilizing elevated non-CpG modification ratios to distinguish cDNA from genomic DNA limits application in species possessing naturally widespread non-CpG methylation. Nonetheless, by overcoming the historic base-level ambiguity of bisulfite sequencing, Joint-Cabernet establishes an enzymatic framework for dissecting how cytosine modifications coordinate mammalian gene regulation across brain maturation, aging, and neurodegenerative disease progression.
Subject of Research: Single-cell multi-omic profiling of DNA methylation, hydroxymethylation, and gene expression in the mammalian brain
Article Title: Joint-Cabernet enables scalable single-cell joint profiling of transcriptomes with DNA methylation and hydroxymethylation in the brain
Article References: Bai, Y., Yuan, T., Ren, L., Huan, Y., Yang, F., Li, Y., Zhang, A., Liu, Y., Tian, T., Kang, N., Xie, X., Wang, Q., Chen, D., Chen, W., Zheng, Y., Xie, X. S., & Cao, Y. (2026). Joint-Cabernet enables scalable single-cell joint profiling of transcriptomes with DNA methylation and hydroxymethylation in the brain. Genome Biology. https://doi.org/10.1186/s13059-026-04351-0
Image Credits: AI Generated
DOI: 10.1186/s13059-026-04351-0
Keywords: Joint-Cabernet, DNA methylation, DNA hydroxymethylation, 5-methylcytosine, 5-hydroxymethylcytosine, single-cell sequencing, transcriptomics, epigenomics, mouse brain, TET dioxygenases, bisulfite-free sequencing, gene regulation, chromatin compartments, neurogenomics, APOBEC3A, MERFISH spatial imputation, differentially hydroxymethylated regions
News Source: Juliet Wilcox. (November 19, 2026). Joint-Cabernet Profiles Mouse Brain Gene Expression Alongside Methylation and Hydroxymethylation. Scienmag.