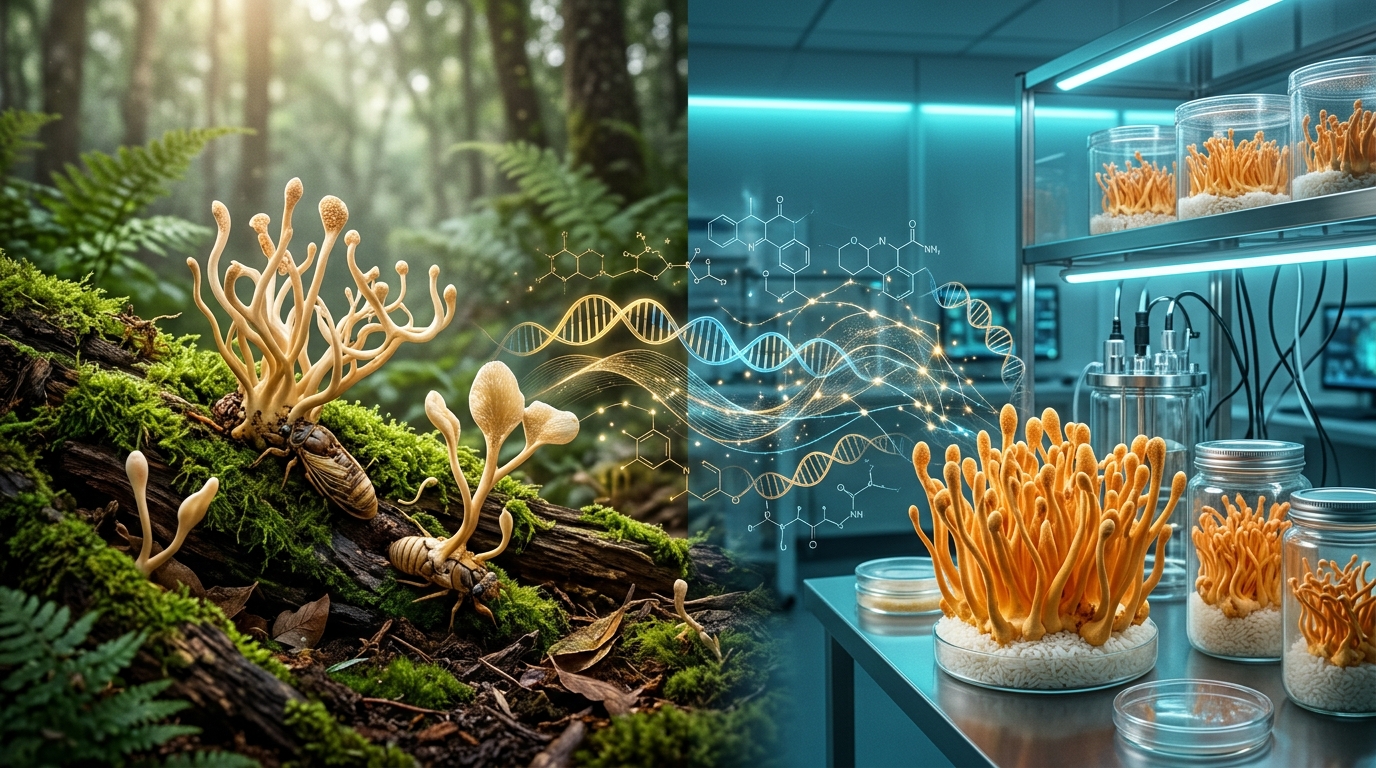
A fungus prized in China as both food and medicine behaves very differently depending on whether it grows in a bamboo forest or on an oat-filled tray in a laboratory. A new multi-omics study of Cordyceps chanhua, an entomopathogenic fungus whose fruiting body was approved as a new food raw material by China’s National Health Commission in 2021, has mapped in unprecedented detail how gene expression and metabolite profiles diverge between wild and cultivated material at two distinct life stages. The findings, published in Food Production, Processing and Nutrition, do not declare one source superior to the other. Instead, they reveal that the molecular choreography underlying development differs fundamentally between the two growth environments, and they identify candidate pathways and metabolites that future cultivation experiments can now test.
The research team, led by Wei Ji of the Lianyungang Academy of Agricultural Sciences and colleagues, compared four sample types: artificially cultivated fruiting bodies (AF) and mycelia (AM) of the cultivar Yunchan No. 1, and wild fruiting bodies (WF) and host-associated sclerotial or mycelial tissues (WM) collected from a hilly bamboo forest in Yixing, Jiangsu Province. Each group contained three independent biological replicates. The cultivated strain was domesticated from wild resources originally collected in the same geographic region and bamboo forest area, yet the authors caution that strain-level genetic equivalence was not confirmed, so all cultivated-wild comparisons must be read as source-associated rather than strain-controlled.
Using transcriptome sequencing, untargeted metabolomics, targeted nucleoside quantification, and integrated transcriptome-metabolome analysis, the researchers identified 1,000 differentially expressed genes and 370 differentially accumulated metabolites in the AF versus AM comparison, and 1,080 differentially expressed genes and 602 differentially accumulated metabolites in the WF versus WM comparison. These numbers indicate extensive stage-associated transcriptional and metabolic remodeling in both systems. In cultivated samples, the differential metabolites were mainly tied to amino acid metabolism, membrane lipid remodeling, purine metabolism, and secondary metabolism. In wild samples, they clustered more closely around glutathione metabolism, central carbon metabolism, and redox regulation, suggesting that natural conditions impose additional demands on cellular stress management and energy balance.
The metabolomic quality control was rigorous. Principal component analysis showed that the first two components explained 39.3 percent and 19.5 percent of total variance, and quality control samples clustered tightly, confirming instrument stability. Chemical classification of all identified metabolites revealed a diverse profile dominated by carboxylic acids and derivatives at 28.43 percent, fatty acyls at 12.61 percent, and prenol lipids at 11.56 percent. When the team screened for differential metabolites between stages, the AF versus AM comparison yielded 196 upregulated and 174 downregulated metabolites, while the WF versus WM comparison produced 327 upregulated and 275 downregulated metabolites. Same-stage source comparisons were even more striking: 607 differential metabolites separated cultivated from wild fruiting bodies, and 1,074 separated cultivated from wild mycelia, indicating that source-associated variation was particularly pronounced at the mycelial stage.
Pathway enrichment analysis added nuance. In cultivated fruiting bodies, significantly enriched KEGG terms included several with animal-derived names such as long-term depression, circadian entrainment, and nicotine addiction. The authors stress that these labels reflect the original KEGG database nomenclature and were interpreted in the fungal context as conserved eukaryotic signaling modules, including cyclic nucleotide signaling, calcium-associated transduction, and light or circadian response components, rather than as animal-specific processes. In wild fruiting bodies, enriched pathways included oxidative phosphorylation, glutathione metabolism, vitamin B6 metabolism, and central carbon metabolism, the latter retained under its original KEGG name but interpreted as conserved glycolysis, tricarboxylic acid cycle, and energy metabolism rather than anything cancer-related.
Targeted quantification of two bioactive nucleosides revealed a clear stage-dependent split. N6-(2-hydroxyethyl)adenosine, or HEA, a characteristic nucleoside-related component of C. chanhua, was significantly enriched in mycelia under both conditions. Cultivated mycelia contained 0.83 plus or minus 0.08 percent HEA, significantly higher than the 0.51 plus or minus 0.08 percent found in cultivated fruiting bodies, and wild mycelia held 0.76 plus or minus 0.05 percent versus 0.49 plus or minus 0.08 percent in wild fruiting bodies, both at P less than 0.01. Adenosine showed the opposite pattern in cultivated material, reaching 1.47 plus or minus 0.15 percent in cultivated fruiting bodies compared with 0.84 plus or minus 0.12 percent in cultivated mycelia, while the wild fruiting body increase was more modest at 0.70 plus or minus 0.07 percent versus 0.56 plus or minus 0.05 percent in wild mycelia. These distinct accumulation patterns suggest that the mycelial stage could be further evaluated as a potential source of HEA-enriched material, while fruiting body cultivation may merit investigation for adenosine-oriented production, though the authors emphasize that such implications require validation through targeted quantification, bioactivity assessment, and process evaluation.
Integrated analysis of gene-metabolite correlations exposed perhaps the most striking difference between the two systems. Cultivated samples showed a more compact and highly connected association network, with most selected gene-metabolite pairs exceeding a Pearson correlation coefficient of 0.8 and forming clear clustering patterns. One gene cluster correlated positively with lysophospholipids, oxidized lipid derivatives, metabolic intermediates such as L-glutamic acid 5-phosphate and adenylosuccinate, and the triterpenoid-related metabolite ganolucidic acid A, while another cluster correlated negatively. Wild samples, by contrast, displayed weaker overall transcriptome-metabolome correlations, with most absolute coefficients below 0.6 and no clear clustering. The wild network was dominated by negative correlations and lacked a single dominant hub node, suggesting a more dispersed, multi-pathway association pattern under natural conditions. The authors interpret this as reflecting greater inter-individual variation and a more complex regulatory background rather than lower regulatory efficiency.
Protein-protein interaction network analysis identified two highly connected candidate transcripts. In the cultivated comparison, TRINITY_DN131_c0_g1, significantly downregulated in fruiting bodies relative to mycelia, carried best-hit annotations related to pheromone-processing carboxypeptidase KexA and serine carboxypeptidase, with the Peptidase_S10 conserved domain. In the wild comparison, TRINITY_DN80_c0_g1, downregulated in wild fruiting bodies, showed annotations related to alpha-glucosidase II or glucan 1,3-alpha-glucosidase and glycosyl hydrolase family 31. Both were interpreted as candidate functional nodes based on de novo transcript assembly, homology-based annotation, and predicted network topology, not as validated key regulators, and neither was confirmed by qRT-PCR in this study.
Several limitations frame the conclusions. The study compared only mycelial and fruiting body stages, omitting primordia and intermediate stages, so results reflect stage-associated differences rather than a continuous developmental trajectory. Developmental and source effects could not be fully disentangled because wild and cultivated samples differed in habitat, substrate or host background, and potentially genetic background. Most metabolites from the untargeted LC-MS dataset were putatively annotated by MS/MS database matching rather than confirmed with authentic standards, so biological interpretation focused on metabolite classes and pathway-level trends rather than individual unvalidated identities. Only HEA and adenosine were independently quantified with reference standards.
Despite these constraints, the study delivers a rich, testable framework for future work. The compact gene-metabolite network in cultivated samples identifies candidate molecular modules associated with developmental processes and target metabolite accumulation, while the dispersed pattern in wild samples suggests that natural heterogeneity may complicate quality standardization. The authors propose that candidate pathways related to purine metabolism, amino acid metabolism, carbohydrate metabolism, membrane lipid remodeling, and developmental regulation could serve as molecular indicators for monitoring bioactive metabolite accumulation and product-quality stability in future controlled cultivation experiments. Because cultivation parameters were not experimentally manipulated here, these associations remain hypotheses rather than validated optimization targets. Controlled experiments involving nutrient supply, temperature, light, and other factors are now needed to determine whether the molecular signatures uncovered in this study can translate into more reliable, higher-quality production of this increasingly important functional food resource.
Subject of Research: Comparative transcriptomic and metabolomic analysis of wild versus cultivated Cordyceps chanhua across developmental stages
Article Title: Integrated transcriptomic and metabolomic analyses reveal developmental and metabolic differences between wild and cultivated Cordyceps chanhua
Article References: Ji, W., Liu, X., Tang, X., Su, W., Chai, W., Ge, J., Hou, L., & Li, F. (2026). Integrated transcriptomic and metabolomic analyses reveal developmental and metabolic differences between wild and cultivated Cordyceps chanhua. Food Production, Processing and Nutrition, 8(1), Article 83. https://doi.org/10.1186/s43014-026-00436-2
Image Credits: AI Generated
DOI: 10.1186/s43014-026-00436-2
Keywords: Cordyceps chanhua, transcriptomics, metabolomics, fruiting body development, HEA, adenosine, nucleoside accumulation, cultivated versus wild, KEGG pathway enrichment, gene-metabolite correlation, entomopathogenic fungus, functional foods
News Source: Juliet Wilcox. (November 4, 2026). Wild and cultivated Cordyceps chanhua show distinct gene and metabolite patterns. Scienmag.