Flibanserin, the first non-hormonal drug approved for hypoactive sexual desire disorder, has long been hampered by an awkward pharmacokinetic problem: only about a third of an oral dose actually reaches the bloodstream, because the liver extensively metabolizes the molecule before it can act. A new study reports a possible workaround. Researchers engineered a flibanserin-loaded cubosomal hydrogel for delivery through the skin, and in a rat model the transdermal formulation produced roughly double the systemic drug exposure of an oral suspension, while slowing absorption in a way the authors argue could suit a condition that requires sustained rather than pulsatile drug levels.
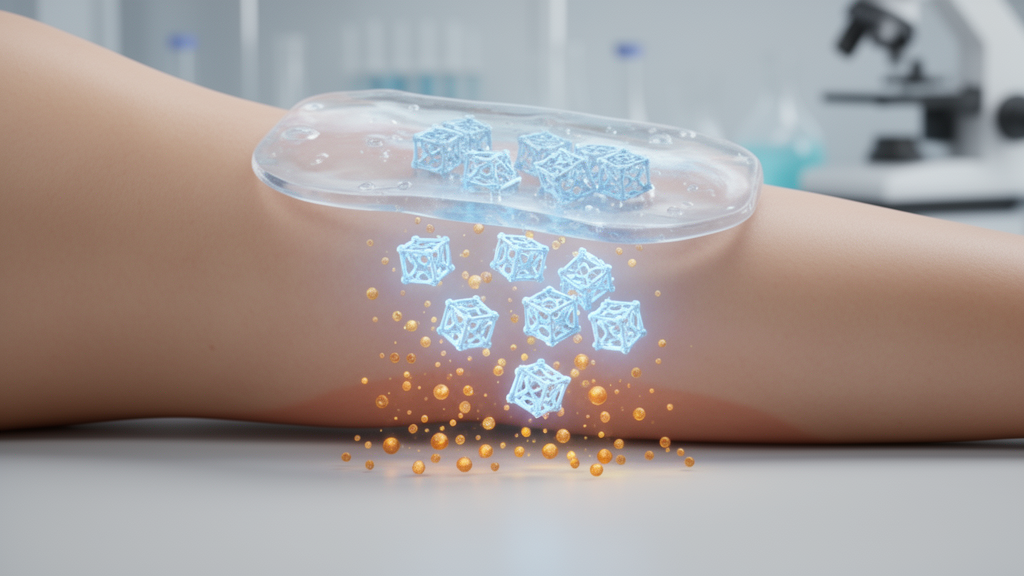
The work, published in Future Journal of Pharmaceutical Sciences, combined a statistical formulation strategy with an in vivo pharmacokinetic comparison. Cubosomes are lipid-based nanocarriers formed when amphiphilic lipids such as monoolein self-assemble in water into highly ordered three-dimensional structures: a single continuous lipid bilayer twisted into a complex spiral that defines two non-intersecting water channels. This bicontinuous cubic architecture can host hydrophobic drugs within the lipid membrane, hydrophilic drugs in the aqueous channels, and amphiphilic drugs at the interface, with loading capacity and stability that the authors note surpasses conventional liposomes. Because their biomimetic lipid composition can interact with the stratum corneum, the skin’s outermost barrier, cubosomes are hypothesized to fuse with and fluidize the skin’s lipid layers, creating transient pathways for drug diffusion while acting as a high-concentration reservoir on the skin surface.
To build the carrier, the team turned to a Box-Behnken design, a response-surface method from design of experiments that generated 17 formulations, including five center points to estimate pure error. Three variables were varied: monoolein content, poloxamer 407 content, and polyvinyl alcohol content, all expressed as percent by weight. Particle size served as the sole optimization response, a choice the researchers justified on the grounds that particle size is a critical quality attribute influencing skin interaction, permeation, physical stability, and subsequent systemic absorption. Prior work suggests particles larger than 600 nanometers tend to remain confined to the upper skin layers, whereas particles of 300 nanometers or less can penetrate more deeply through the skin strata.
The prepared formulations showed particle sizes ranging from 245 to 493 nanometers. Analysis of variance confirmed the data fit a two-factor interaction model with an R-squared of 0.9931, a highly significant F-value of 190.82, and a nonsignificant lack of fit. Both monoolein and polyvinyl alcohol exerted significant inverse effects on particle size, and all three interaction terms were significant, pointing to complex interdependencies among the formulation variables rather than simple additive effects. The authors interpret the monoolein effect as promoting a more well-defined hydrophobic domain that favors self-assembly of smaller, thermodynamically stable particles, while polyvinyl alcohol acts as a steric stabilizer that adsorbs at the lipid-water interface and suppresses aggregation. Notably, this monoolein finding diverges from some earlier cubosome studies that reported the opposite trend, a discrepancy the authors attribute to differences in surfactant systems or homogenization energy.
Numerical optimization identified an ideal composition of 9.93 percent monoolein, 1.36 percent poloxamer 407, and 0.03 percent polyvinyl alcohol. When prepared, this formulation yielded an observed particle size of 234.3 plus or minus 8.6 nanometers, closely matching the model-predicted value of 236.8 nanometers with a percentage error of just 1.07 percent. Entrapment efficiency reached 83.2 plus or minus 5.7 percent, indicating that shrinking the particles did not compromise the carrier’s capacity to hold the lipophilic drug, which the authors link to the hydrophobic effect driving amphiphile self-assembly and the protected reservoir formed within the lipid bilayers. Transmission electron microscopy confirmed lyotropic liquid crystalline nanoparticles with a bicontinuous cubic structure.
The optimized nanodispersion was then incorporated into a hydrogel matrix of 2 percent hydroxypropyl methylcellulose K4M, chosen for its compatibility with lipid nanocarriers and its ability to provide topical-appropriate viscosity while supporting sustained release. The resulting cubogel was white, uniform, and homogeneous, with no grittiness, a viscosity of 2.15 plus or minus 0.19 Pa.s, which falls within the 2 to 3 Pa.s range reported as optimal for dermatological gels, and a spreadability diameter of 4.7 plus or minus 0.21 centimeters, within the ideal 4 to 6 centimeter range for topical products. Compared with conventional transdermal patches, the authors note, a gel offers greater dosage flexibility, reduced potential for skin irritation, and better cosmetic acceptability.
The decisive test was pharmacokinetic. Twenty-four male Wistar rats were randomly divided into two groups of twelve. One group received the optimized cubogel transdermally on a shaved dorsal skin area of roughly four square centimeters at a flibanserin dose of 10 milligrams per kilogram; the other received a raw flibanserin suspension orally at the same dose. The authors specify that male rats were used to enable a controlled comparison of formulation-dependent pharmacokinetic parameters without potential estrous-cycle-related effects on systemic exposure, not to evaluate therapeutic efficacy in a disease model. Plasma flibanserin concentrations were measured with a validated LC-MS/MS method showing excellent linearity, and the parameters were derived by non-compartmental analysis.
The transdermal route clearly outperformed oral dosing on exposure measures. The area under the plasma concentration-time curve extrapolated to infinity was 130.7 plus or minus 21.3 nanogram-hours per milliliter after transdermal application versus 66.4 plus or minus 4.3 after oral administration, a statistically significant difference corresponding to a relative bioavailability of 196.8 percent, or a 1.97-fold increase in systemic exposure. Time to peak concentration lengthened from 0.5 to 2 hours, and mean residence time extended from 4.2 to 6.3 hours, indicating prolonged absorption. Peak plasma concentration was numerically lower after transdermal dosing, 19.1 versus 21.4 nanograms per milliliter, but the difference was not statistically significant. The authors attribute the gains to avoidance of gastrointestinal and hepatic first-pass metabolism together with prolonged skin residence and the permeation-enhancing properties of the cubosomal hydrogel, and they suggest the flattened profile could minimize peak-related side effects while maintaining therapeutic concentrations.
The researchers are careful to frame these results as an initial, formulation-focused validation rather than clinical proof. The study was conducted entirely in male rats, and the authors acknowledge that future studies in female animals, with appropriate consideration of estrous-cycle stage, will be needed to strengthen translational relevance for women with HSDD. They also note that in vitro release, ex vivo skin permeation, permeation flux, lag time, permeability coefficient, and skin deposition studies using Franz diffusion cells were not performed and would provide additional mechanistic insight into the respective contributions of monoolein, poloxamer 407, and the cubosomal nanostructure. Comprehensive long-term and accelerated stability studies, dermal safety assessment, chronic toxicity testing, and pharmacological efficacy evaluation are all listed as necessary next steps before the platform could advance toward patients.
Even with those caveats, the study offers a coherent demonstration of a design chain running from statistical optimization through nanostructure characterization to in vivo performance. The authors argue that the clinical value of the cubogel derives not from any single parameter but from the deliberate engineering linking particle size control, nanostructural integrity, and hydrogel rheology to a sustained, first-pass-independent pharmacokinetic profile. They further propose the approach as a transferable framework for other poorly bioavailable, lipophilic drugs that could similarly benefit from cubosome-in-hydrogel transdermal delivery, positioning particle-size-directed design of experiments as a generalizable starting point for engineering such systems. For a drug whose oral formulation has long underserved the patients it was approved to help, that framework, if it survives the remaining preclinical hurdles, could matter well beyond flibanserin itself.
Subject of Research: A cubosomal transdermal hydrogel formulated to improve the delivery of flibanserin for hypoactive sexual desire disorder.
Article Title: Transdermal flibanserin cubosomal hydrogel for hypoactive sexual desire disorder: box–behnken optimization and pharmacokinetic profiling in a rat model
Article References: Badr-Eldin, S. M., Aldawsari, H. M., Abdelahkeem, E., Abualsunun, W., Eshmawi, B. A., Ahmed, O. A. A., Kotta, S., & Naguib, M. J. (2026). Transdermal flibanserin cubosomal hydrogel for hypoactive sexual desire disorder: box–behnken optimization and pharmacokinetic profiling in a rat model. Future Journal of Pharmaceutical Sciences, 12(1), Article 86. https://doi.org/10.1186/s43094-026-01018-2
Image Credits: AI Generated
DOI: 10.1186/s43094-026-01018-2
Keywords: flibanserin, cubosomes, transdermal delivery, hypoactive sexual desire disorder, nanotechnology, pharmacokinetics, Box-Behnken design, hydrogel, drug bioavailability, rat model, HPMC, first-pass metabolism
News Source: Ophelia Keating. (October 29, 2026). Nano Cubosome Gel Boosts Flibanserin Skin Delivery Nearly Twofold in Rats. Scienmag.