Selective breeding that incorporates ruminal microbial gene profiles can substantially reduce enteric methane emissions in beef cattle while circumventing severe performance penalties in growth and feed efficiency, according to a genomic modeling investigation conducted at Scotland’s Rural College. Enteric fermentation within ruminant livestock accounts for approximately seventy-eight percent of the agrifood sector’s 5.28 gigatonnes of carbon dioxide equivalent emitted as methane, a greenhouse gas possessing twenty-seven to thirty times the warming potential of carbon dioxide over a century-long horizon. In addition to driving global atmospheric warming, enteric methanogenesis forfeits up to twelve percent of total dietary gross energy consumed by the animal, representing an intrinsic biological inefficiency. Because direct quantification of enteric methane output requires specialized and costly monitoring infrastructure that impedes continuous herd-scale phenotyping, the research team developed a microbiome-driven breeding methodology. Their framework identifies specific microbial functional genes in the rumen that accurately forecast host genetic merit for methane production, offering a practical avenue for cost-effective genomic mitigation.
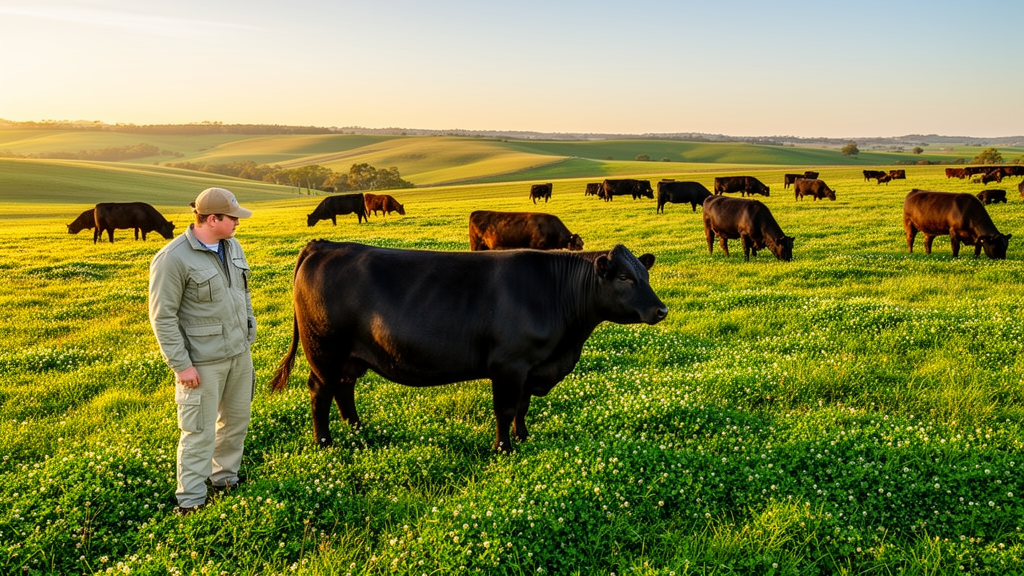
The study evaluated a population of 359 beef steers representing purebred Luing, Charolais-cross, and rotational Aberdeen Angus-Limousin crosses fed two basal diets with forage-to-concentrate ratios of 480:520 and 80:920 grams per kilogram on a dry matter basis. Across the five experimental years spanning 2011 to 2017, performance testing under uniform husbandry conditions yielded an average daily gain of 1.476 kilograms, daily feed intake of 11.04 kilograms, cold carcass weight of 372.9 kilograms, and a feed conversion ratio of 7.61 kilograms of feed per kilogram of gain. Daily methane production, monitored continuously during the final forty-eight hours of a seventy-two-hour stay in indirect open-circuit respiration chambers across 287 steers, averaged 190.5 grams per day with a standard deviation of 52.8 grams per day. This methane output demonstrated substantial phenotypic variability, exhibiting a coefficient of variation of 0.28, which surpassed the relative variation observed in average daily gain, daily feed intake, and cold carcass weight.
Genomic parameter estimation utilizing Bayesian univariate and bivariate analyses revealed that daily methane production in this controlled finishing cohort possessed a high host heritability of 0.56, with a ninety-five percent highest posterior density interval ranging from 0.26 to 0.86. Daily feed intake exhibited an estimated heritability of 0.61, while average daily gain and cold carcass weight displayed heritabilities of 0.39 and 0.48, respectively. Genetic correlations among all measured production metrics and daily methane production were markedly positive, uncovering deeply unfavorable biological associations for conventional mitigation. Daily feed intake exhibited an exceptionally strong genetic correlation of 0.88 with methane production, alongside genetic correlations of 0.93 with average daily gain and 0.84 with carcass weight. Direct growth measures also showed unfavorable genetic correlations with daily methane output, estimated at 0.58 for average daily gain and 0.61 for cold carcass weight, whereas phenotypic correlations were consistently below 0.40.
To explore microbiome architecture without relying on physical gas enclosures, researchers sequenced postmortem ruminal digesta collected at slaughter across 359 steers, generating between 26 and 159 million paired reads per animal. The bioinformatics pipeline aligned assembled contigs to the Kyoto Encyclopedia of Genes and Genomes database, identifying 7,976 functional microbial genes that were filtered to 3,632 genes present in at least seventy percent of samples at relative abundances exceeding 0.001 percent. Following additive log-ratio transformation using ribulose-phosphate 3-epimerase as the invariant reference denominator, the team performed 3,631 bivariate analyses against methane output. They identified a prioritized subset of forty-three microbial genes, designated MG43, that exhibited posterior probabilities exceeding eighty percent of possessing non-zero genetic correlations with methane production, phenotypic correlations exceeding 0.10, and individual expected correlated responses above two percent of the population mean.
Functional annotation of the forty-three candidate genes revealed precise biochemical mechanisms driving hydrogen disposal and methanogenesis in the ruminal ecosystem. Positively correlated genes predominantly coordinated oxidoreductase processes, including the rnfD gene encoding a subunit of ferredoxin:NAD+ oxidoreductase, lldE encoding L-lactate dehydrogenase, and oorD encoding 2-oxoglutarate ferredoxin oxidoreductase, which supply reduced electron donors such as NADH and reduced ferredoxin to methanogenic archaea. The nicotinamide mononucleotide transporter K03811 exhibited the single strongest genetic association with methane, while genes governing coenzyme A biosynthesis such as menI and PANK1_2_3 supported the production of volatile fatty acids, specifically acetate and butyrate, which generate molecular hydrogen during fermentation. Conversely, microbial genes negatively associated with methane output orchestrated essential maintenance, structural resilience, amino acid metabolism, and thermal adaptation, including the serA, metA, and cas2 genes, reflecting non-methanogenic biomass growth that serves as an effective hydrogen sink.
Evaluating genomic estimated breeding values across several selection scenarios demonstrated the efficacy of microbiome-driven breeding. Selecting the top ten percent of the herd solely on measured methane in a univariate framework achieved an estimated mitigation response of seventeen percent per generation. When a higher selection intensity of 2.06 was applied, selection targeting these forty-three microbial genes delivered a 23.25 percent reduction in methane output. Integrating direct chamber measurements alongside the forty-three microbial genes pushed expected mitigation to 26.83 percent per generation, establishing that functional metagenomic markers provide predictive accuracy equal or superior to physical gas quantification.
However, univariate genomic selection targeted purely at lowering methane production produced pronounced collateral deterioration across economically vital performance traits. Selecting the best ten percent of animals solely for reduced methane caused a severe correlated decline of 9.67 percent in average daily gain, a 4.93 percent reduction in cold carcass weight, and a 10.70 percent drop in daily feed intake under estimated genetic correlations. When direct methane measurements were combined with production traits and the forty-three microbial genes, expected methane mitigation reached 24.22 percent at a selection intensity of 1.76, but average daily gain plunged by 11.67 percent. Even in sensitivity analyses that replaced estimated genetic correlations with lower phenotypic values, selecting directly for lower methane still induced an unfavorable 2.95 percent decline in daily gain, confirming that unconstrained selection against methane simply depresses animal appetite and overall growth.
To circumvent these deleterious production penalties, the researchers formulated two complementary selection strategies that sever the adverse link between greenhouse gas abatement and herd productivity. First, they derived genomic estimated breeding values for residual methane production, defined mathematically as the genetic residual of daily methane output adjusted for average daily gain and cold carcass weight using partial genetic regression coefficients. Selecting the top ten percent of animals for lower residual methane achieved a meaningful 13.68 percent reduction in methane production while keeping correlated changes in daily gain and carcass weight near zero.
As an alternative biological approach, the investigators formulated a second microbiome-driven model, termed MDB41, centered on forty-one distinct microbial genes that displayed simultaneous favorable genetic correlations with both reduced methane and enhanced average daily gain. Although these genes had relaxed selection thresholds, requiring posterior probabilities greater than sixty percent and correlated responses exceeding one percent of each trait mean, they encompassed pathways supporting carbohydrate hydrolysis, nucleotide synthesis, and environmental sensing, such as amyA, afcA, and ciaH. When deployed in a separate evaluation model without production traits, MDB41 achieved an expected methane reduction of fourteen percent per generation at a selection intensity of 1.755 while preserving average daily gain with a small positive trajectory. Crucially, integrating MDB41 into a multivariate model alongside recorded production traits re-introduced an adverse 10.79 percent decline in daily gain, demonstrating that MDB41 must operate independently to avoid overriding its beneficial microbial balance.
Scatter plots of individual breeding values confirmed the presence of correlation-breaking animals within the population that combined low residual methane or low residual feed intake with superior genetic potential for weight gain and carcass yield, exceeding a five percent dual improvement. While practical implementation must address the logistical demands of gastric tube sampling and high-throughput sequencing pipelines, longitudinal evidence shows that a single ruminal sample taken during the growing-finishing period provides stable predictive utility over time. The authors emphasize that observed selection responses reflect intensive single-herd testing under respiration chamber conditions and will naturally moderate in commercial herds undergoing multi-trait multi-objective selection. Nevertheless, by harnessing host-regulated ruminal microbial genes and residual genetic criteria, livestock industries can deploy cost-effective genomic selection that mitigates enteric methane emissions while safeguarding beef cattle productivity.
Subject of Research: Microbiome-driven genomic selection for enteric methane mitigation in beef cattle
Article Title: Optimisation of selection for methane mitigation by integrating production traits with ruminal microbiome-driven breeding in beef cattle
Article References: Nguyen, T. Q., Martínez-Álvaro, M., Lima, J., Gorjanc, G., Cleveland, M. A., & Roehe, R. (2026). Optimisation of selection for methane mitigation by integrating production traits with ruminal microbiome-driven breeding in beef cattle. Genetics Selection Evolution, 58(1), Article 47. https://doi.org/10.1186/s12711-026-01053-w
Image Credits: AI Generated
DOI: 10.1186/s12711-026-01053-w
Keywords: Beef Cattle, Enteric Methane, Methane Mitigation, Rumen Microbiome, Microbial Genes, Microbiome-Driven Breeding, Genomic Selection, Genomic Estimated Breeding Values, Residual Methane Production, Average Daily Gain, Daily Feed Intake, Cold Carcass Weight, Residual Feed Intake, Respiration Chambers, Methanogenesis, Scotland's Rural College
News Source: Morgan Morrow. (November 13, 2026). Microbiome-Driven Breeding Strategy Cuts Beef Cattle Methane Without Impairing Growth. Scienmag.