Researchers have demonstrated that the AlphaFold3 (AF3) artificial intelligence framework provides superior structural prediction accuracy for adeno-associated virus (AAV) capsid protein subunits compared to established tools like RoseTTAFold2 (RF2) and ESMFold. Adeno-associated viruses are widely utilized in gene therapy as delivery vectors, and understanding the multimeric organization of their outer capsid shell is essential for rational engineering. By benchmarking these AI models against high-resolution experimental structures obtained through cryo-electron microscopy and X-ray crystallography, the study evaluated the accuracy of predicted monomeric VP1, VP2, and VP3 subunits and local multimeric interfaces across ten AAV serotypes.
The study found that AF3 consistently produced lower Cα root-mean-square deviation (RMSD) values—a metric of structural divergence from experimental references—across all three VP classes. For the primary benchmarking of VP3 subunits, which constitute the majority of the capsid shell, AF3 achieved a mean RMSD of 0.535 ± 0.152 Å. This performance was significantly better than the RF2 mean RMSD of 0.917 ± 0.050 Å and the ESMFold mean RMSD of 0.982 ± 0.076 Å. Statistical analysis using a Friedman test confirmed significant differences in structural accuracy among the methods, with post hoc Wilcoxon signed-rank tests indicating that AF3 significantly reduced the paired median RMSD compared to both RF2 and ESMFold.
In addition to structural accuracy, AF3 exhibited a reliable confidence-accuracy association for the VP3 monomer panel. The study identified a significant negative correlation between the predicted local distance difference test (pLDDT) score and observed backbone RMSD (Spearman ρ = −0.745, p = 0.013), suggesting that higher predicted confidence levels accurately correspond to lower structural error. Conversely, the RF2 and ESMFold models did not display statistically significant confidence-accuracy associations in this dataset. This ability to integrate lower structural error with predictive confidence markers suggests that AF3 offers a robust computational foundation for analyzing the known AAV fold family.
The research also extended to the multimeric benchmarking of VP3-only assemblies, focusing on the 2-fold dimeric and 3-fold trimeric interfaces critical for capsid symmetry and stability. AF3 produced low RMSD values over the aligned regions across all serotypes, maintaining structural fidelity within these complex assemblies. In paired dimer comparisons, AF3 demonstrated significantly lower RMSD than classical docking frameworks, including RosettaDock, HADDOCK, and LZerD. Similar results were observed for trimer comparisons, where AF3 consistently yielded lower backbone RMSD values than the specialized docking approaches evaluated in the study.
While classical docking tools like RosettaDock and HADDOCK have historically been used to sample rigid-body arrangements for viral capsids, the study highlighted significant variability in their performance. RosettaDock often showed low contact recovery as measured by the fraction of native contacts, while HADDOCK output cluster quality varied considerably across different serotypes and restraint configurations. The LZerD approach, which utilized statistical potentials such as GOAP and DFIRE to select representative models, also exhibited higher median RMSD values compared to AF3. These findings indicate that while diverse computational routes exist for modeling subunit arrangements, the generative diffusion framework implemented in AF3 provides a high degree of consistency.
The study included full-length VP1 and VP2 sequences to evaluate the reconstruction of the conserved capsid core in the presence of N-terminal extensions that are known to be flexible and often unresolved in experimental structural data. By excluding these flexible regions from quantitative RMSD scoring, the researchers successfully compared the prediction of the common core architecture. Qualitative inspection suggested that the AF3-predicted N-terminal extensions were often directed toward the capsid interior, a positioning that aligns with structural hypotheses regarding internal domain organization proposed in studies of other AAV serotypes.
The benchmark analysis addressed potential training-data overlap by evaluating the availability of experimental reference structures relative to AF3’s published structural-training cutoff of 30 September 2021. Although some reference structures became available in 2022–2023, exploratory comparisons grouped by reference date did not show statistically significant differences in structural accuracy within the study’s panels. This framing emphasizes that the results reflect performance as structural benchmarking within an experimentally characterized fold family, rather than isolated inference on unseen targets.
Structural modeling remains a critical component of rational AAV engineering, where capsid modification strategies such as surface loop insertion, receptor retargeting, and epitope masking depend on accurate interface definitions. Local geometric differences in subunit packing can influence the biological function and production efficiency of engineered vectors. By providing a comparative assessment of current AI-driven modeling workflows, the study highlights how these computational tools may support the in silico pre-screening of capsid candidates before progressing to experimental validation.
Limitations of the study include the reliance on specific server-default workflows for the AI models and representative-model selection criteria for the docking suites. While the benchmark provides a comprehensive snapshot of performance across ten serotypes, the research acknowledges that controlled ablation experiments or matched-input configurations would be required to fully isolate the contributions of factors such as MSA-derived features, language-model embeddings, and symmetry-aware restraints. Furthermore, the findings are specific to the characterized AAV serotypes included, and the transferability of these structural models to highly diversified, synthetic, or ML-designed capsid variants remains a target for future prospective assessment.
In conclusion, the study establishes AF3 as a highly accurate structural modeling approach for AAV capsid subunits and symmetry interfaces. By consistently achieving the lowest backbone RMSD and providing meaningful confidence-accuracy associations, AF3 facilitates the interpretation of known structural features within the AAV family. As the field moves toward large-scale computational design, these structural predictions offer a valuable resource for guiding rational engineering efforts and understanding the relationship between capsid structure and vector function.
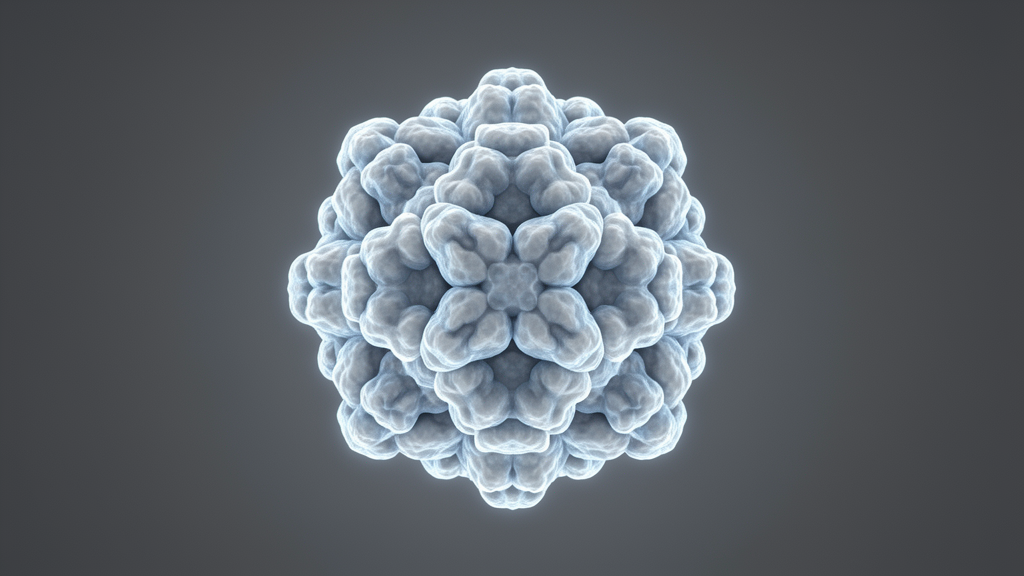
Biological delivery using adeno-associated viruses relies on a sixty-subunit icosahedral capsid assembled from VP1, VP2, and VP3 proteins in an approximate 1:1:10 ratio. While all three share a conserved C-terminal core corresponding to VP3, the extended N-termini of VP1 and VP2 carry specialized functional machinery. Specifically, the basic BR3 cluster within the shared N-terminal domain participates in nuclear translocation, whereas the VP1-unique extension houses a phospholipase A2 sequence that mediates endosomal escape. Because structural ordering in experimentally resolved capsids is concentrated within the VP3 segment, comparing full-length VP1 and VP2 models against experimental VP3 coordinates enabled researchers to evaluate whether deep-learning architectures could reconstruct the conserved structural core when presented with differing sequence contexts.
Across the full-length monomer evaluations, AlphaFold3 maintained consistent accuracy regardless of subunit length. For VP1 subunits, AlphaFold3 achieved a mean Cα RMSD of 0.491 ± 0.101 Å, outperforming RoseTTAFold2 at 0.925 ± 0.058 Å and ESMFold at 1.037 ± 0.086 Å. For VP2, AlphaFold3 averaged 0.486 ± 0.100 Å, compared with 0.901 ± 0.050 Å for RoseTTAFold2 and 0.983 ± 0.089 Å for ESMFold. Exploratory Friedman tests assessing sequence context across VP classes showed no statistically significant differences in structural deviation across the three subunits for AlphaFold3 (χ2 = 0.200, p = 0.905) or RoseTTAFold2 (χ2 = 1.800, p = 0.407). In contrast, ESMFold exhibited significant variability across subunit types (χ2 = 6.000, unadjusted p = 0.0498), driven by elevated deviations in the longer VP1 sequences. Local model confidence reflected similar patterns, with AlphaFold3 attaining mean VP1 pLDDT scores of 84.5 ± 0.6 and VP2 scores of 89.5 ± 2.4.
Detailed quantitative benchmarking of multimeric assemblies further demonstrated AlphaFold3’s capacity to model local quaternary architecture. In paired evaluations of two-fold dimeric interfaces across serotypes, AlphaFold3 significantly reduced median paired RMSD by 0.2895 Å compared to HADDOCK (95% bootstrap confidence interval: −0.3795 to −0.1810 Å), by 0.2950 Å compared to LZerD (95% CI: −0.3145 to −0.2260 Å), and by 0.2535 Å compared to RosettaDock (95% CI: −0.4970 to −0.1405 Å). After Benjamini–Hochberg false discovery rate correction, all dimer comparisons were statistically significant at q = 0.000977. Similarly, in three-fold trimer assemblies, AlphaFold3 reduced median paired RMSD by 0.3705 Å relative to HADDOCK (q = 0.003906), by 0.3175 Å relative to LZerD (q = 0.002930), and by 0.1680 Å relative to RosettaDock (q = 0.003906). Across individual serotypes, AlphaFold3 trimer deviations ranged from 0.225 Å in AAV1 to 0.732 Å in AAV2, with corresponding interface predicted template modeling scores spanning 0.71 to 0.79.
The comparative docking workflows relied on distinct sampling principles and restraint formulations that influenced multimer outcomes. For HADDOCK, docking incorporated ambiguous interaction restraints derived from conserved capsid stability hotspots, including the five-residue segment mapped from AAV2 (residues M604, V605, D608, R609, and D610) to corresponding positions in other serotypes. Although selected HADDOCK clusters achieved local RMSD values below 1.0 Å for specific serotypes, alternative clusters and runs produced large deviations exceeding 10 to 40 Å. For LZerD, candidate poses were scored using geometric Zernike descriptors combined with GOAP, DFIRE, and ITScore statistical potentials. While the selected LZerD models yielded bounded local interface deviations between 0.722 and 1.042 Å for trimers, expanding these assemblies into hexamers revealed rotational misorientations relative to the experimental crystallographic symmetry axes, demonstrating that local backbone agreement does not guarantee correct global subunit orientation.
These structural benchmarks provide crucial context for machine-learning-guided vector design platforms that generate extensive sequence diversity. Earlier studies demonstrated that deep-learning models could generate over 110,000 viable capsid variants across diversified sequence libraries. Accurate structural modeling of local subunit interfaces establishes a physical foundation to interpret such candidate substitutions before conducting resource-intensive vector production experiments.
Subject of Research: Benchmarking AI-based protein structure prediction for AAV capsid engineering
Article Title: Benchmarking AI structure prediction tools for rational AAV vector engineering
Article References: Zhang, W., Liu, Y., Li, J., Noriega, H. A., & Wang, X. S. (2026). Benchmarking AI structure prediction tools for rational AAV vector engineering. Frontiers in Drug Discovery, 6, 1908034. https://doi.org/10.3389/fddsv.2026.1908034
Image Credits: AI Generated
DOI: 10.3389/fddsv.2026.1908034
Keywords: AlphaFold3, AAV, structural biology, gene delivery, capsid engineering, protein structure prediction, RoseTTAFold2, ESMFold, cryo-EM, viral vectors, bioinformatics, computational modeling, AI in drug discovery, multimeric assembly, protein-protein interaction, AAV serotypes, RMSD analysis, rational design
News Source: Kristina Jarvis. (November 16, 2026). AlphaFold3 Outperforms Conventional Tools in Predicting Adeno-Associated Virus Capsid Structures. Scienmag.