A newly identified homozygous splice-site variant in the PGAP1 gene has been reported in a patient with neurodevelopmental impairment, according to an original article published in Molecular Biology Reports on 25 October 2026. The variant, designated c.1173+1G>A, was absent from all public genetic databases and was classified as likely pathogenic under American College of Medical Genetics and Genomics (ACMG) guidelines, adding a new entry to the short list of PGAP1 variants documented worldwide.
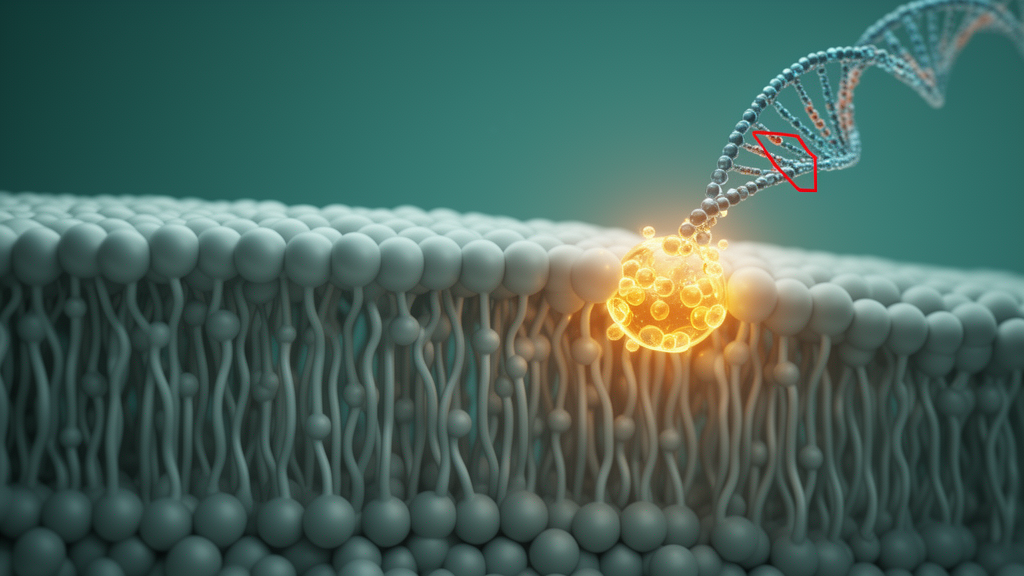
PGAP1 encodes an inositol deacylase located in the endoplasmic reticulum, where it carries out the first step of remodeling glycosylphosphatidylinositol (GPI) anchors. Genetic defects in this pathway form a recently recognized group of autosomal recessive neurodevelopmental disorders with highly variable clinical presentations.
Pathogenic variants in PGAP1 have so far been reported in only a limited number of families globally. Affected individuals described in the literature show features including global developmental delay, intellectual disability, seizures, microcephaly, and variable systemic manifestations. The new report contributes both a novel variant and an expanded picture of the phenotypic range associated with this gene.
The affected patient presented with neurodevelopmental impairment and carried the splice-site variant in homozygous form, meaning both copies of the gene carried the same change.
Because the study relied on computational prediction rather than direct RNA or protein analysis, the authors assessed the variant’s likely effects using multiple in silico tools. SpliceAI, Human Splicing Finder, and varSEAK were used to evaluate the potential impact on splicing, and these analyses consistently predicted significant disruption of normal splicing at the affected site.
To explore possible structural consequences, the researchers used AlphaFold for protein structure prediction and PyMOL for visualization. Their modeling suggested conformational alterations associated with a predicted in-frame deletion in the protein, providing a plausible mechanistic link between the splice defect and loss of the deacylase’s normal function, though such predictions remain computational estimates rather than experimentally confirmed outcomes.
Segregation analysis was performed to establish the inheritance pattern within the family and to supply additional evidence for the variant’s classification. The homozygous state of the variant in the affected patient, combined with its absence from public databases and the consistent computational predictions, supported the likely pathogenic designation under ACMG criteria.
In addition to the case description, the authors conducted a comprehensive review of previously reported PGAP1 cases to broaden the known phenotypic spectrum of the disorder.
The report’s conclusions carry two practical implications. First, they expand the phenotypic heterogeneity attributed to PGAP1-related disorders and add molecular detail to the understanding of GPI-anchor remodeling defects.
The findings are limited by their basis in a single patient, computational rather than experimental functional validation, and the small number of PGAP1 cases available for comparison.
Subject of Research: A novel PGAP1 splice-site variant associated with neurodevelopmental impairment in a GPI-anchor remodeling disorder
Article Title: A novel PGAP1 splice-site variant expanding the clinical and molecular spectrum of GPI-anchor disorders
Article References: Hashemi, S.-S., Moazzami, R., & Miri, A. (2026). A novel PGAP1 splice-site variant expanding the clinical and molecular spectrum of GPI-anchor disorders. Molecular Biology Reports, 53(1), Article 1823. https://doi.org/10.1007/s11033-026-13043-8
Image Credits: AI Generated
DOI: 10.1007/s11033-026-13043-8
Keywords: PGAP1, GPI-anchor, splice-site variant, neurodevelopmental disorder, autosomal recessive, in silico splicing prediction, ACMG classification, global developmental delay, intellectual disability, genomic diagnosis, endoplasmic reticulum, inositol deacylase
News Source: Juliet Wilcox. (October 29, 2026). Novel PGAP1 Splice-Site Variant Broadens Known Spectrum of GPI-Anchor Disorders. Scienmag.