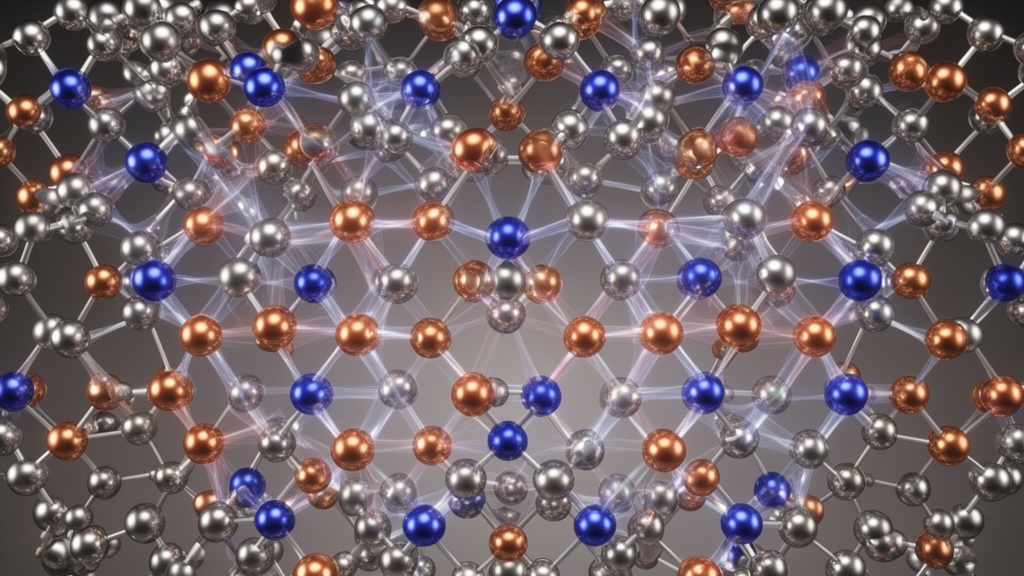
A comprehensive scientific review published in the Journal of Materials Science demonstrates that machine learning interatomic potentials have emerged as a transformative computational framework capable of overcoming longstanding scale limitations in the atomistic modeling of high-entropy alloys. High-entropy alloys, which contain multiple principal elements in non-dilute concentrations, exhibit remarkable physical, mechanical, and functional properties governed by heterogeneous local chemical environments and extensive configurational diversity. Traditional modeling has relied extensively on first-principles density functional theory calculations to evaluate thermodynamic, mechanical, magnetic, and defect properties. However, because density functional theory computational costs scale approximately cubically with the number of atoms, practical first-principles calculations are restricted to cells of a few hundred atoms and molecular dynamics trajectories of tens of picoseconds. The review, authored by researchers including Yuji Ikeda, Xiang Xu, Pranav Kumar, Jörg Neugebauer, Li-Fang Zhu, Xi Zhang, Fritz Körmann, and Blazej Grabowski, establishes that machine learning interatomic potentials successfully bridge this critical gap, providing near-quantum accuracy while dramatically reducing computational overhead to allow realistic simulations across millions of atoms.
The central difficulty in investigating high-entropy alloys stems from their combinatorially vast compositional and configurational spaces. Experimental evaluations frequently measure macroscopic properties that average over intricate microstructures and diverse local atomic environments, obscuring the precise atomic-scale mechanisms responsible for observed phenomena such as cryogenic fracture toughness or high-temperature strength retention. While density functional theory offers parameter-free electronic insights, its strict length and time scale boundaries prevent direct sampling of statistical fluctuations, extended defect structures, and slow kinetic processes. Machine learning interatomic potentials, trained on rigorously constructed electronic structure datasets, reproduce density functional theory energies and forces several orders of magnitude faster. This computational acceleration enables large-scale molecular dynamics simulations tracking systems of ten to the sixth power atoms over nanosecond durations, and in specialized implementations reaching up to ten to the eighth power atoms. As a consequence, researchers can directly simulate complex physical mechanisms including chemical short-range order, dislocation motion, solute drag, grain boundary segregation, and hydrogen absorption in compositionally complex materials.
Constructing reliable machine learning potentials for multi-component alloys introduces distinct methodological hurdles, particularly because the number of interatomic combinations scales sharply with the number of constituent elements. In standard many-body representations, the parameter space grows combinatorially with the species count, demanding innovative model architectures. To resolve this problem, modern formalisms implement chemical embedding strategies that map discrete elemental species onto continuous, low-dimensional latent vectors. In one prominent application, researchers compressed twenty-five transition metals into an artificial neural network potential using only three or four pseudo-chemical dimensions without degrading accuracy, and successfully interpolated values for unrepresented elements like rhenium and osmium. Complementing embedding techniques, low-rank tensor factorizations applied to density-based representations, such as multi-atomic cluster expansions and graph-based atomic cluster expansions, successfully eliminate combinatorial parameter scaling. Furthermore, active learning frameworks systematically explore configuration space using iterative workflows, identifying structural uncertainties through ensemble committee disagreement, Bayesian predictive deviations, or D-optimality matrix selection algorithms to efficiently direct supplemental electronic structure calculations.
Sampling extended structural defects represents an equally demanding task, as defects like dislocation cores, crack tips, and grain boundaries govern mechanical deformation but often exceed the cell sizes accessible to direct quantum calculations. Naive extraction of finite atomic clusters introduces artificial surfaces and truncates long-range interactions, which severely degrades potential accuracy. Advanced reconstruction protocols resolve this limitation by applying rotational and mirror symmetry to reconstruct periodic dislocation dipoles or by embedding extracted clusters into periodic supercells with surrounding vacuum regions repopulated via grand-canonical Monte Carlo steps. An alternative conceptual strategy relies on decomposing complex defect structures into constituent planar defects, as demonstrated by sequential active learning used to capture Kear-Wilsdorf locks in nickel-aluminum compounds. Additionally, one-shot sampling frameworks generate diverse small-cell structures spanning various space groups, successfully leveraging the fundamental locality of machine learning potentials to accurately describe general grain boundaries without requiring explicit large-scale grain boundary configurations during initial training.
Accurate mechanical and thermodynamic modeling also demands training datasets that encompass chemically ordered and segregated environments rather than relying exclusively on ideal random solid solutions. Density functional theory investigations have frequently represented random alloys through special quasirandom structures that maximize configurational entropy. However, experimental observations from transmission electron microscopy and atom probe tomography confirm that high-entropy alloys frequently develop chemical short-range order. To model these ordering phenomena, researchers utilize on-lattice machine learning interatomic potentials, notably low-rank potentials employing tensor train decompositions. By training on relaxed atomic structures and spin-polarized electronic data, low-rank potentials implicitly capture local lattice distortions and magnetic interactions. In chromium-cobalt-nickel alloys, accounting for the lowest energy magnetic state during potential training accurately predicted order-disorder transitions yielding face-centered cubic ordered phases consistent with experimental measurements, whereas models trained on non-spin-polarized data incorrectly yielded molybdenum-platinum-type ordering, highlighting the decisive influence of magnetism on chemical arrangement.
While computationally efficient on-lattice models capture static configurational energies, they cannot account for finite-temperature vibrational coupling with local chemical order. To address this limitation, off-lattice machine learning potentials are integrated into hybrid Monte Carlo and molecular dynamics simulations, as well as replica exchange molecular dynamics. In these hybrid workflows, periodic Monte Carlo species swaps rapidly equilibrate chemical configurations, while concurrent molecular dynamics trajectories permit atomic coordinates to relax and explore thermal vibrations. Studies utilizing these techniques reveal that chemical short-range order significantly modifies stacking fault energies, defect formation kinetics, and dislocation glide resistance. Furthermore, evaluations of local order demonstrate that traditional Warren-Cowley parameters, which measure only pair correlations within nearest neighbor shells, are insufficient to uniquely define chemical configurations. Distinct local arrangements exhibiting identical Warren-Cowley parameters can exhibit substantially different Kullback-Leibler divergences and cohesive energies. Additionally, simulations incorporating athermal forced atomic swaps demonstrate that non-equilibrium processing, such as rapid solidification or work hardening, produces remnant steady-state ordering configurations distinct from any thermal equilibrium state.
Machine learning potentials also provide powerful avenues for computing temperature-dependent vibrational contributions to Gibbs energies beyond standard quasi-harmonic approximations. At elevated temperatures approaching melting thresholds, anharmonic lattice vibrations contribute significantly to free energies and phase stability. Traditional thermodynamic integration requiring continuous ab initio molecular dynamics trajectories across multiple coupling parameters demands prohibitive computational resources. By introducing accurate machine learning potentials as intermediate reference models, researchers perform two-stage integration or direct upsampling via free-energy perturbation. In equiatomic vanadium-chromium-tantalum-tungsten alloys, direct upsampling from machine learning potentials to density functional theory achieved free-energy agreement within one and a half millielectronvolts per atom without requiring ab initio molecular dynamics. Similar implementations in face-centered cubic vanadium-cobalt-nickel improved order-disorder transition temperature predictions by approximately one third. Beyond bulk stability, machine learning potentials combined with solid-liquid coexistence simulations and free-energy workflows accurately determine melting temperatures, fusion enthalpies, and volume changes across refractory alloys, providing essential quantitative data for thermodynamic Calphad databases.
The review details significant progress in clarifying point-defect energetics and diffusion kinetics in compositionally complex alloys. Unlike pure metals, where point-defect formation energies are single-valued, high-entropy alloys exhibit broad energy distributions driven by local chemical variations. Thermodynamically consistent evaluations require referencing defect chemical potentials to the multi-component alloy rather than unary phases, which can be accomplished using the Widom-type substitution technique or average vacancy formation relations. Investigations in chromium-iron-nickel and aluminum-scandium-titanium-zirconium-hafnium alloys show that vacancy formation energies depend strongly on first nearest neighbor chemical compositions and atomic arrangements, while second nearest neighbors exert negligible effects. Integrating these distributed formation energies produces effective temperature-dependent formation Gibbs energies and negative effective formation entropies, leading to pronounced low-temperature deviations from classic linear Arrhenius behavior. Machine learning potentials extended to diffusion transition states have similarly unlocked temperature-dependent migration barriers, capturing complex self-diffusion pathways and anisotropic kinetic behaviors.
At the mesoscale, machine learning potentials provide vital insights into extended defects, plastic deformation, and fracture mechanisms. In body-centered cubic niobium-molybdenum-tantalum-tungsten, simulations revealed a reduced mobility difference between screw and edge dislocations compared with constituent unary metals, pointing to an active role for edge dislocations in plastic strain. Furthermore, atomistic simulations identified a cross-slip locking mechanism that temporarily immobilizes screw dislocations through cross-kink and jog formation, producing substantial solid-solution strengthening. Chemical short-range order exerts a dual effect on plasticity: it elevates the energy barrier for kink-pair nucleation along screw dislocations while simultaneously reducing solute drag on edge dislocations. In polycrystalline simulations exceeding one hundred million atoms, machine learning potentials capture grain boundary segregation, showing that elements such as niobium preferentially accumulate at boundaries while tungsten concentrates in grains. Under mechanical loading, plastic deformation initiates via dislocation emission from grain boundaries, with grain boundary stability and local short-range order dictating whether an alloy undergoes brittle intergranular fracture or extensive boundary-mediated plasticity.
Functional applications of high-entropy alloys, particularly solid-state hydrogen storage, represent another emerging frontier for machine learning potentials. Certain high-entropy compositions, including titanium-vanadium-zirconium-niobium-hafnium, absorb hydrogen up to hydrogen-to-metal ratios of two and a half, undergoing phase transformations to face-centered cubic structures with hydrogen occupying both tetrahedral and octahedral interstitial sites. In complex hexagonal C14 and cubic C15 Laves phases, interstitial hydrogen exhibits distinct site preferences among symmetrically inequivalent coordination environments. Moment tensor potentials applied to titanium-chromium Laves hydrides successfully reproduced preferential occupation of specific tetrahedral sites and captured temperature-dependent hydride stability. Moreover, path-integral molecular dynamics enabled by machine learning potentials revealed that quantum tunneling effects substantially lower hydrogen migration barriers at room temperature. Integrating these atomistic insights into data-driven machine learning models enables the identification of predictive descriptors, such as valence electron concentration and local binding energy distributions, to guide the accelerated design of reversible room-temperature storage alloys.
The latest technological shift centers on universal machine learning interatomic potentials, or foundation models, pre-trained on massive datasets spanning tens of thousands to hundreds of millions of diverse chemical configurations. Architectures such as graph-based cluster expansions, equivariant message-passing networks, and related foundation frameworks allow rapid exploration across the periodic table without requiring de novo training sets for every alloy. Benchmark assessments on graphical processing units demonstrate that universal models can achieve energy accuracies matching system-specific potentials, particularly when trained on chemically diverse datasets constructed via entropy-maximization protocols. Nevertheless, current foundation models face important challenges, including potential energy surface softening caused by ground-state data bias, high computational inference costs, and the risk of catastrophic forgetting during task-specific fine-tuning. Developing knowledge-distillation strategies to compress foundation models into lightweight student architectures, alongside incorporating explicit magnetic moments and atomic charge degrees of freedom, will be critical to fully establishing machine learning interatomic potentials as predictive engines for advanced alloy design.
Subject of Research: Atomistic simulations of high-entropy alloys using machine learning interatomic potentials
Article Title: Atomistic simulations of high-entropy alloys: from density functional theory to machine-learning interatomic potentials
Article References: Ikeda, Y., Xu, X., Kumar, P., Neugebauer, J., Zhu, L.-F., Zhang, X., Körmann, F., & Grabowski, B. (2026). Atomistic simulations of high-entropy alloys: from density functional theory to machine-learning interatomic potentials. Journal of Materials Science. https://doi.org/10.1007/s10853-026-13520-2
Image Credits: AI Generated
DOI: 10.1007/s10853-026-13520-2
Keywords: High-Entropy Alloys, Machine Learning Interatomic Potentials, Density Functional Theory, Chemical Short-Range Order, Molecular Dynamics, Thermodynamic Integration, Direct Upsampling, Point Defects, Dislocation Dynamics, Grain Boundary Segregation, Universal Foundation Models, Active Learning, Chemical Embedding, Hydrogen Storage Materials, Free Energy Perturbation, Warren-Cowley Parameters, Phase Stability
News Source: Teresa Odom. (November 16, 2026). Machine Learning Interatomic Potentials Overcome Density Functional Theory Limits in Complex Alloys. Scienmag.